Unveiling proton-regulated Mo=O/Mo-O- centers in reconstructed Ni2Mo-NiOOH/MoOx interfaces toward accelerated proton-coupled electron transfer for urea oxidation reaction
 0
Abstract
Urea electrolysis for hydrogen production offers the dual benefits of low energy consumption and pollutant degradation; however, the sluggish proton-coupled electron transfer (PCET) kinetics remain a critical bottleneck limiting performance enhancement. To overcome this challenge, we herein construct an in situ reconstructed NiMo alloy electrocatalyst featuring highly efficient proton-capturing sites to accelerate reaction kinetics. Under anodic conditions, the NiMo alloy undergoes surface reconstruction to form a Ni2Mo-NiOOH/MoOx composite structure, where NiOOH functions as the oxidative active site driving the urea oxidation reaction (UOR), while Mo=O/Mo-O- species in MoOx act as proton-capturing centers. These sites rapidly sequester protons released during the dehydrogenation of *-NH2 intermediates, substantially lowering the energy barrier of the key reaction step and facilitating the PCET process. Simultaneously, Mo-induced charge redistribution optimizes the adsorption of reaction intermediates, enabling synergistic modulation of electron and proton transfer. Benefiting from these mechanisms, the as-fabricated catalyst requires only 1.341 V to achieve a current density of 100 mA·cm-2, exhibiting outstanding reaction kinetics. Moreover, a urea degradation efficiency of 97.21% is attained within 420 min, demonstrating the synergistic enhancement of energy conversion and pollutant removal. This work not only elucidates the pivotal role of PCET modulation in UOR via efficient proton-capturing sites but also provides a new strategy for the rational design of high-performance electrocatalytic systems.
Keywords
INTRODUCTION
In recent years, driven by the global energy structure transition and the “dual carbon” goals, the development of efficient and sustainable hydrogen production technologies has emerged as a major research focus in the energy field. Among them, water electrolysis for hydrogen production is regarded as one of the key pathways to realizing the hydrogen economy, owing to its inherent advantages of being green, clean, and highly compatible with renewable energy[1-4]. However, conventional water electrolysis systems rely heavily on high-purity freshwater resources, and against the backdrop of growing global water scarcity, this limitation severely hinders their large-scale deployment[5-7]. Therefore, developing alternative anodic reactions to replace the conventional oxygen evolution reaction (OER) is of great significance for reducing energy consumption and expanding the application scenarios of electrolysis systems.
Against this background, the urea oxidation reaction (UOR) has attracted widespread attention due to its unique advantages[8-14]. Conventional water electrolysis consists of the cathodic hydrogen evolution reaction (HER) and the anodic OER, among which OER involves a complex four-electron transfer process with sluggish reaction kinetics, acting as the critical bottleneck limiting the overall energy conversion efficiency[15-22]. In contrast, the theoretical potential of UOR is only 0.37 V, far lower than the 1.23 V of OER, which can significantly reduce the energy consumption of electrolysis[23-26]. Furthermore, urea is widely present in industrial wastewater, agricultural aquaculture wastewater, and municipal domestic sewage. Its electrocatalytic oxidation enables the synergistic realization of hydrogen production and pollutant degradation, thereby constructing an integrated system for energy conversion and environmental remediation[27-30]. From an environmental perspective, untreated urea-containing wastewater is prone to hydrolysis in natural environments, generating ammonia and its derivatives that in turn trigger water eutrophication and ecological pollution issues[31]. In electrocatalytic systems, by contrast, urea can be selectively oxidized to harmless N2 and CO2, achieving efficient pollutant removal[32-34]. Meanwhile, this reaction process avoids the safety hazards associated with the mixing of H2 and O2 in conventional water electrolysis. Consequently, UOR not only exhibits prominent advantages in energy conversion but also demonstrates significant application potential in environmental governance.
Although UOR exhibits distinct thermodynamic superiority, its reaction pathway involves six-electron transfer[35,36], C-N bond cleavage[37,38] and successive N-H[39,40] dehydrogenation processes. The complicated reaction route leads to sluggish kinetics, making the actual operating potential far higher than the theoretical value[41,42]. Current research mainly concentrates on regulating the adsorption strength of catalysts toward reaction intermediates (e.g., *NH2, *NH, *NCO) to optimize the reaction pathway and lower the energy barrier[43]. Nevertheless, most of these strategies are merely limited to the modulation of electronic structure, while paying insufficient attention to the critical proton transfer behavior during the reaction. In fact, electron and proton transfer are generally coupled in many electrochemical reaction processes through the proton-coupled electron transfer (PCET) mechanism, which plays an important role in multi-step UOR reactions[44,45]. In particular, the conversion of *-NH2 to *-NH belongs to a typical dehydrogenation step. Without efficient proton-accepting sites, the released H+ cannot be transferred timely, which substantially increases the reaction energy barrier and induces intermediate accumulation, thereby restricting the overall reaction rate[39]. Accordingly, starting from the intrinsic reaction nature, constructing catalytic interfaces that can effectively promote proton transfer is a critical route to break the kinetic bottleneck of UOR.
Among diverse catalytic systems, Ni-based materials are recognized as the core active centers for UOR, because they can be in situ converted into highly active NiOOH species under alkaline conditions[46]. The high-valent Ni3+/Ni4+ species in NiOOH can efficiently accelerate the oxidation process of urea molecules. However, single-component Ni-based systems still have apparent limitations. Their relatively high d-band center endows them with strong adsorption capacity toward reaction intermediates. Although this is beneficial to reaction initiation, it easily causes difficult desorption of intermediates, further leading to active site poisoning and restricted reaction kinetics[47]. To overcome the above shortcomings, researchers have proposed introducing secondary metals via an alloying strategy to regulate the electronic structure of Ni. For instance, NiFe[48] and NiCo[49] systems can optimize intermediate adsorption behavior and improve catalytic performance to a certain degree. Nevertheless, these strategies mainly focus on electronic structure regulation and adsorption optimization, and lack targeted design for the pivotal proton transfer step in UOR. Hence, it is still difficult to fundamentally break through the kinetic limitations of the reaction.
Based on the above background, this work proposes a catalytic design strategy via synergistic regulation of active sites and proton-capturing sites. Under electrochemical conditions, the Ni2Mo alloy is further in situ reconstructed into a Ni2Mo-NiOOH/MoOx composite structure, where NiOOH acts as the primary oxidative active site to drive UOR[46,50]. More importantly, the Mo=O/Mo-O- species in the in situ generated MoOx establish efficient proton-capturing sites, which can rapidly capture the H+ released during the dehydrogenation of *-NH2. This significantly reduces the energy barrier of proton transfer and accelerates the key PCET step. This strategy realizes the synergistic modulation of electron and proton transfer, thereby effectively overcoming the bottleneck of restricted dehydrogenation kinetics in conventional catalytic systems. In conclusion, by constructing high-efficiency proton-capturing sites, this study fundamentally resolves the limitation of the critical dehydrogenation step in UOR from the mechanistic perspective, providing a new design insight for the regulation of proton-coupled processes in electrocatalytic reactions.
EXPERIMENTAL
Chemicals and reagents
Nickel sulfate hexahydrate (NiSO4·6H2O), concentrated hydrochloric acid (HCl), and potassium hydroxide (KOH) were purchased from Tianjin Damao Chemical Reagent Factory with analytical reagent (AR) grade. Sodium molybdate dihydrate (Na2MoO4·2H2O) and urea (CO(NH2)2) were obtained from Tianjin Guangfu Technology Development Co., Ltd. Sodium citrate (Na3C6H5O7) was supplied by Tianjin Aqiluopu Trading Co., Ltd., and anhydrous ethanol (C2H6O) was purchased from Tianjin Huihang Chemical Technology Co., Ltd. All the above reagents were used directly without further purification. Nickel foam (NF) with a thickness of approximately 1.5 mm was acquired from Tianjin Anuohe New Energy Technology Co., Ltd. and pretreated before use. In addition, sufficient deionized water was used throughout the entire experimental process.
Preparation of Ni2Mo/NF
4 mmol NiSO4·6H2O, 2 mmol Na2MoO4·2H2O and 4.3 mmol Na3C6H5O7 were separately dissolved in 50 mL deionized water, followed by magnetic stirring for 30 min. Afterward, the pH value of the mixed solution was adjusted to 9 using ammonia solution. Electrodeposition was performed at 40 °C for 10 min under a current density of -180 mA·cm-2, with NF as the working electrode, graphite rod as the counter electrode, and saturated calomel electrode (SCE) as the reference electrode. The obtained electrode was rinsed three times with deionized water and then dried in a vacuum drying oven at 60 °C for 6 h. The as-prepared Ni2Mo/NF catalytic electrode was finally sealed and stored for subsequent use.
Preparation of Ni2Mo-NiOOH/MoOx via in situ reconstruction
In the electrolyte containing 1 M KOH and 0.33 M urea, in situ reconstruction was conducted at a current density of 100 mA·cm-2 for 60 min. Ni2Mo/NF served as the working electrode, graphite rod as the counter electrode, and SCE as the reference electrode. After reconstruction, the sample was ultrasonically treated and dried in an oven to obtain the final product of Ni2Mo-NiOOH/MoOx.
Detailed information on material characterizations and electrochemical measurements is provided in the Experimental Section of the Supporting Information.
RESULTS AND DISCUSSION
Morphological and composition analysis
To clarify the correlation between morphological evolution and phase structure of in situ reconstructed NiMo-based electrocatalysts, systematic scanning electron microscopy (SEM) and transmission electron microscopy (TEM) characterizations were conducted on the as-prepared catalytic electrodes. As shown in Figure 1A1 and A2, the control sample Ni-NiOOH exhibits a uniform nanoparticle array morphology with a compact and flat substrate surface. In comparison, the Ni2Mo alloy [Figure 1B1 and B2] displays a denser particle stacking structure and presents typical alloy substrate features under low-magnification SEM. Further TEM characterization [Supplementary Figure 1] confirms its dense bulk morphology via bright-field TEM observation. The distinct lattice fringes with an interplanar spacing of 0.217 nm in high-resolution transmission electron microscopy (HRTEM) correspond to the (111) crystal plane of Ni2Mo/NF, demonstrating the successful construction of highly crystalline Ni2Mo/NF. The polycrystalline ring patterns in selected area electron diffraction (SAED) further validate the crystallinity of the precursor, which provides a fundamental structural basis for the subsequent anodic reconstruction. After anodic in situ reconstruction, SEM images of Ni2Mo-NiOOH/MoOx [Figure 1C1 and C2] show that the original granular morphology disappears and transforms into a rough, defect-enriched reconstructed layer. The inhomogeneous roughening of the substrate surface directly evidences the drastic surface reconstruction during anodic oxidation. Meanwhile, the introduction of Mo induces surface reconstruction and defect exposure, which not only enlarges the electrolyte contact area and increases the density of active sites, but also creates open channels for subsequent proton transfer.
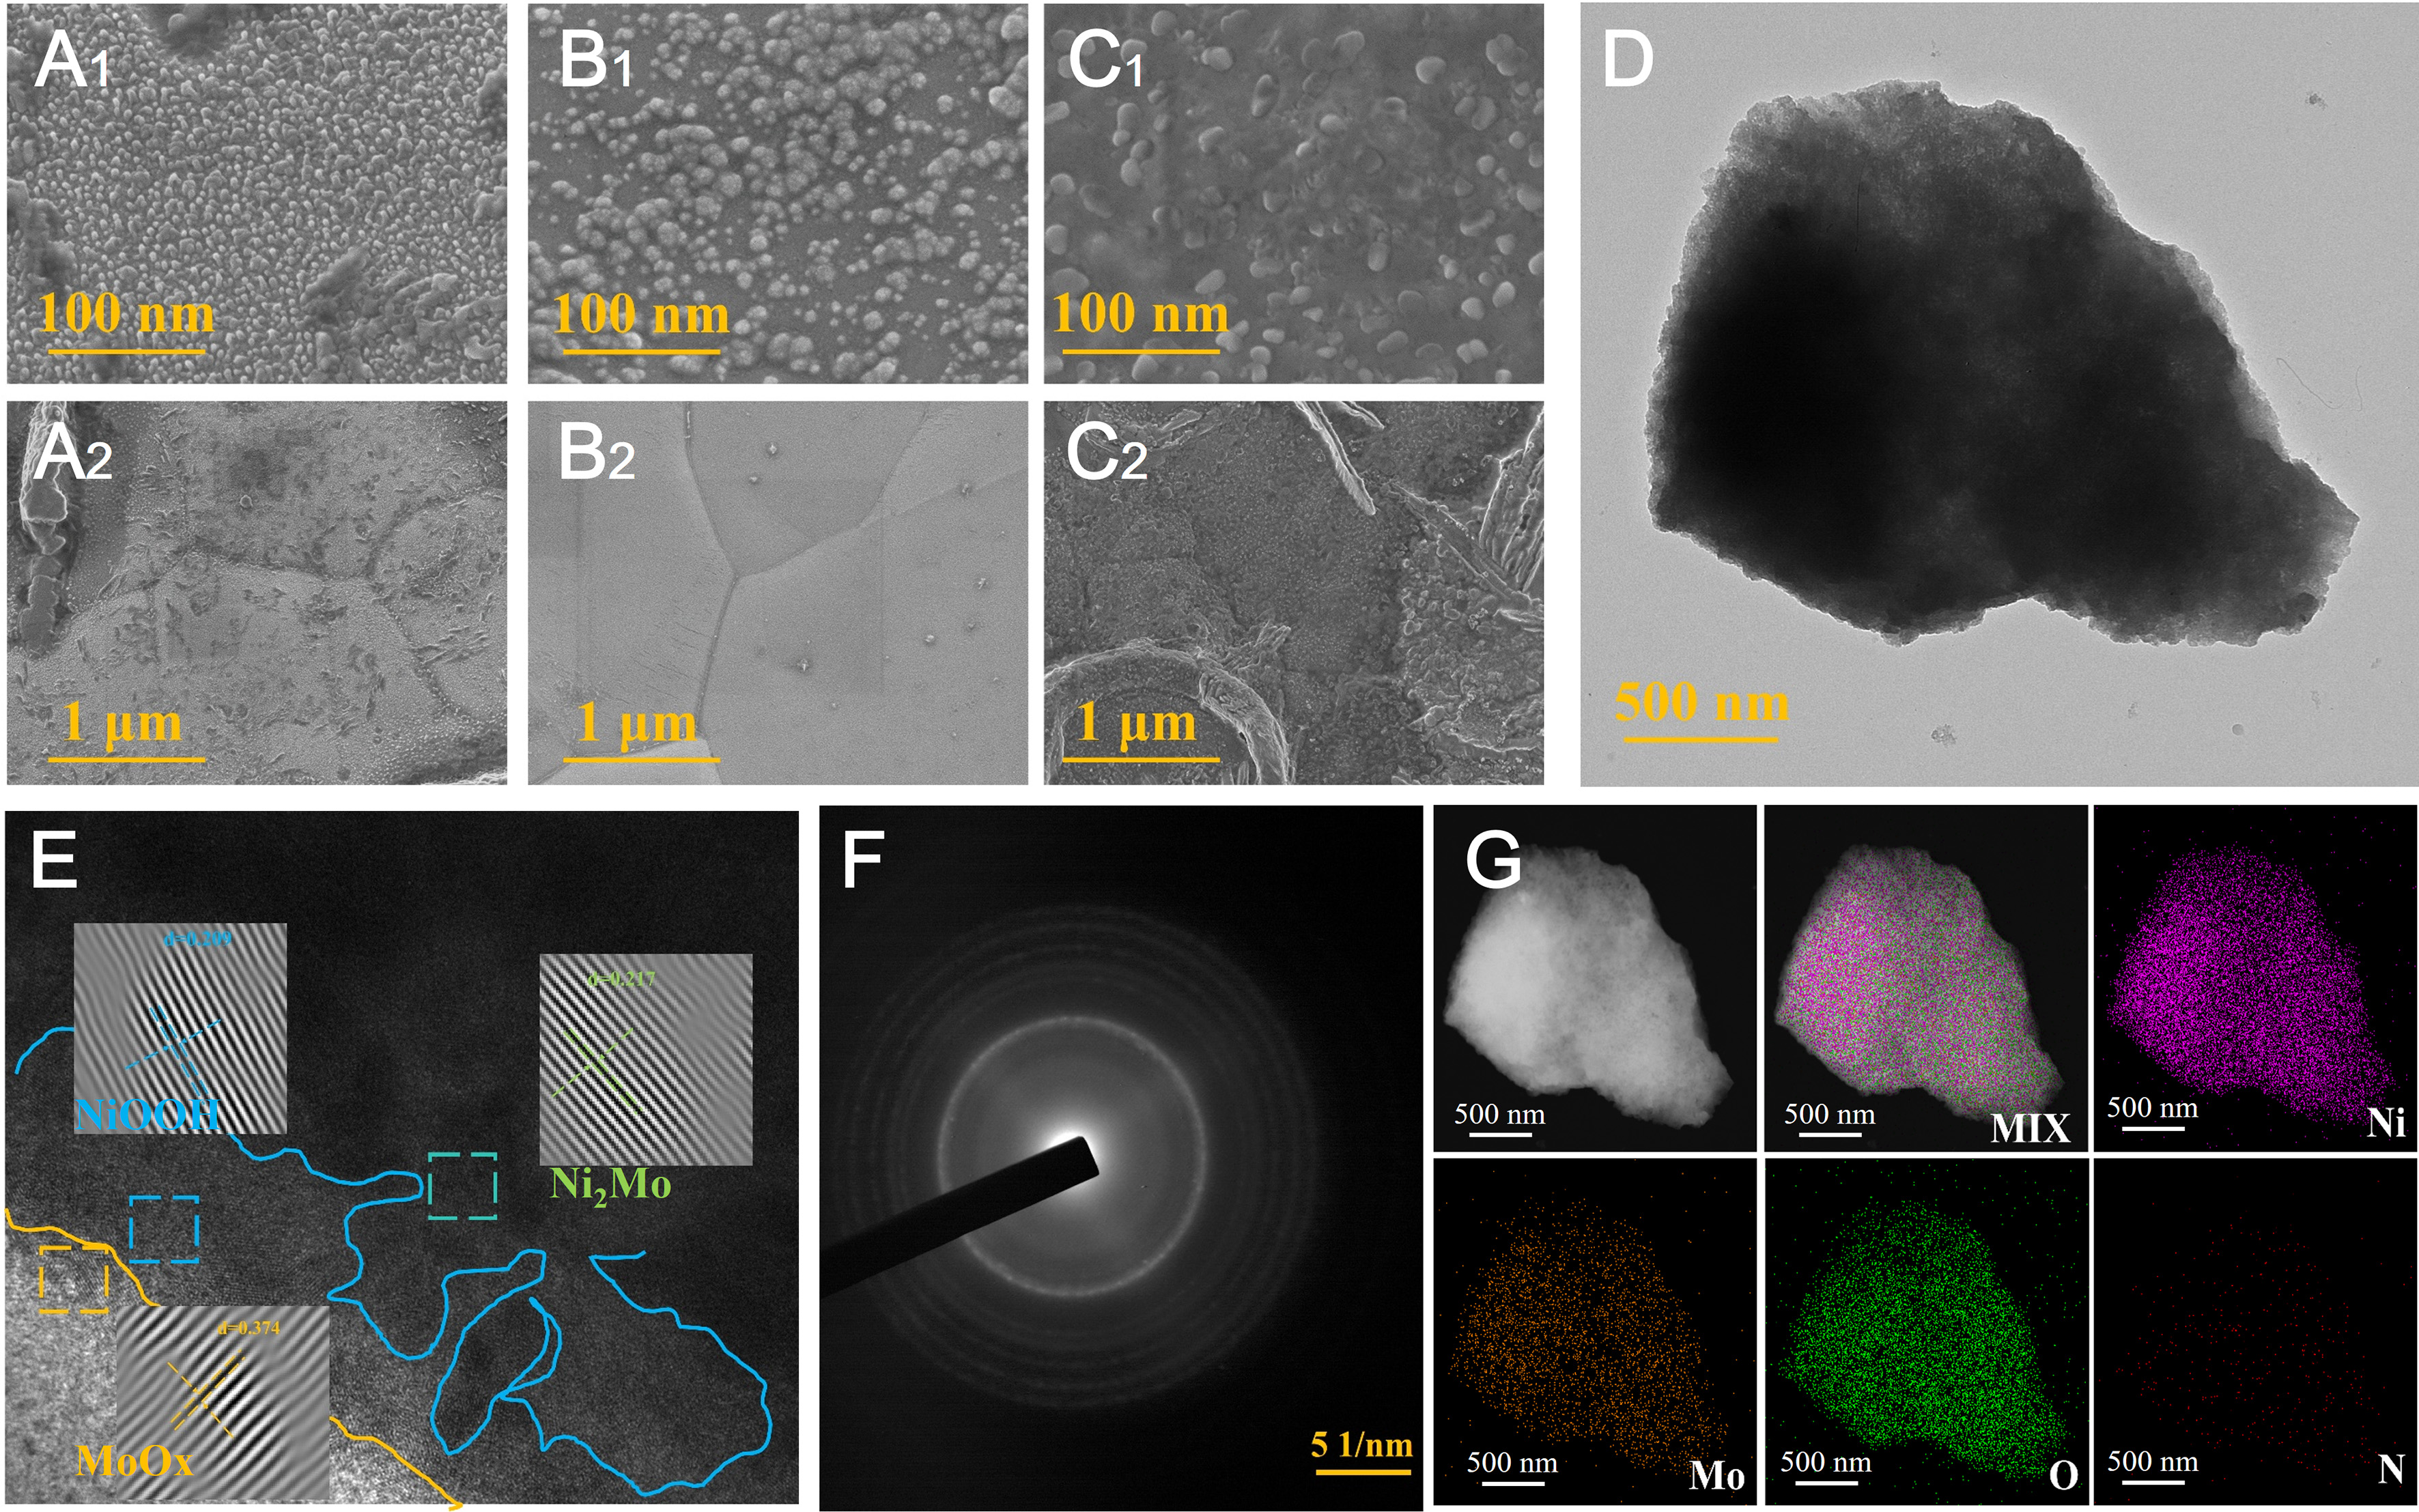
Figure 1. Morphology and structural characterization of the as-prepared catalysts. (A1 and A2) SEM images of Ni-NiOOH; (B1 and B2) SEM images of Ni2Mo/NF; (C1 and C2) SEM images of Ni2Mo-NiOOH/MoOx; (D) TEM image of Ni2Mo-NiOOH/MoOx; (E) HRTEM image of Ni2Mo-NiOOH/MoOx; (F) SAED pattern of Ni2Mo-NiOOH/MoOx; (G) EDS elemental mapping images of Ni2Mo-NiOOH/MoOx (Ni, Mo, O, N). SEM: Scanning electron microscopy; NF: nickel foam; HRTEM: high-resolution transmission electron microscopy; EDS: energy-dispersive X-ray spectroscopy; SAED: selected area electron diffraction.
TEM images of Ni2Mo-NiOOH/MoOx [Figure 1D] reveal that the overall morphology is well preserved after reconstruction, while a thin and translucent reconstructed shell layer emerges at the edge, which is identified as an amorphous/low-crystalline layer. This indicates that the reconstruction process only occurs on the material surface, and the inner core maintains the Ni2Mo/NF phase. Such a core-shell structure provides rapid pathways for charge transport. HRTEM images [Figure 1E] clearly identify a triphase heterostructure, including the amorphous MoOx phase (d = 0.374 nm), crystalline NiOOH active phase (d = 0.209 nm), and Ni2Mo/NF core (d = 0.217 nm). This result directly confirms the formation of Ni2Mo-NiOOH/MoOx heterostructure after in situ reconstruction. The intimate contact of the triphase interfaces builds a favorable microenvironment for the synergistic transfer of electrons and protons. The coexistence of crystalline polycrystalline rings and amorphous halos in the SAED pattern [Figure 1F] further verifies the crystalline-amorphous composite structure. Such amorphous features are conducive to enhancing the dynamic surface reconstruction ability and regulating the adsorption behavior of reaction intermediates. Moreover, energy-dispersive X-ray spectroscopy (EDS) elemental mapping [Figure 1G] confirms the coexistence and distribution of Ni, Mo, O and N elements in the reconstructed catalyst. Although the signals of Mo and N are relatively weak due to their low content and surface-related distribution, they can still be detected throughout the material. The presence of Mo indicates the formation of MoOx during the reconstruction process, while the N signal is mainly attributed to the adsorption of residual urea molecules and N-containing intermediates on the catalyst surface during UOR. This endows abundant Mo=O/Mo-O- proton-capturing sites for UOR. The above characterization results verify that the in situ constructed NiOOH active sites are closely interfaced with MoOx proton-capturing sites, laying a solid structural foundation for the electrocatalytic reaction.
The phase composition and crystal structure of the samples were systematically analyzed via X-ray diffraction (XRD) [Figure 2A]. The characteristic diffraction peaks of the Ni2Mo/NF catalytic electrode located at 43.77°, 50.69°, and 73.65° correspond to the (111), (200), and (220) crystal planes, respectively, which are highly consistent with the standard card of NiMo (PDF#03-2714), confirming the successful preparation of Ni2Mo/NF. The diffraction peaks of the control sample Ni-NiOOH match well with the standard card of β-NiOOH (PDF#06-0141), providing a benchmark for the phase identification of the reconstructed phase. After anodic in situ reconstruction, the XRD pattern of Ni2Mo-NiOOH/MoOx exhibits obvious phase evolution. The intensity of the characteristic peaks belonging to the Ni2Mo precursor decreases significantly, indicating that partial reconstruction of the alloy phase occurs during anodic oxidation. Meanwhile, new characteristic peaks emerge at 37.28° and 66.76°, which are assigned to the (002) and (110) crystal planes of β-NiOOH and perfectly match PDF#06-0141. This result verifies the generation of crystalline NiOOH active phases during the reconstruction process. Combined with TEM analysis, the newly formed MoOx presents an amorphous structure; thus, no crystalline diffraction peaks of MoOx are detected. Fourier transform infrared spectroscopy (FT-IR) spectroscopy was employed to elucidate the evolution of chemical bonds and functional groups on the catalyst surface [Figure 2B]. Ni2Mo/NF is dominated by the metallic state with weak signals of metal-oxygen bonds and hydroxyl functional groups. After anodic in situ reconstruction, Ni2Mo-NiOOH/MoOx shows remarkably enhanced characteristic absorption peaks in the range of 400-800 cm-1, which are attributed to the Ni-O bonds of NiOOH as well as the Mo-O and Mo=O bonds of MoOx species. This confirms the successful formation of NiOOH active phases and MoOx components after reconstruction. Additionally, the stretching vibration intensity of hydroxyl O-H groups at 3,200-3,600 cm-1 increases obviously, suggesting abundant enrichment of hydroxyl groups on the catalyst surface. Furthermore, the rich Mo=O/Mo-O- functional groups in MoOx can serve as efficient proton-capturing sites. Meanwhile, the hydrogen bond network constructed by surface *OH effectively connects proton transport pathways, accelerating the migration rate of H+ released from the dehydrogenation of
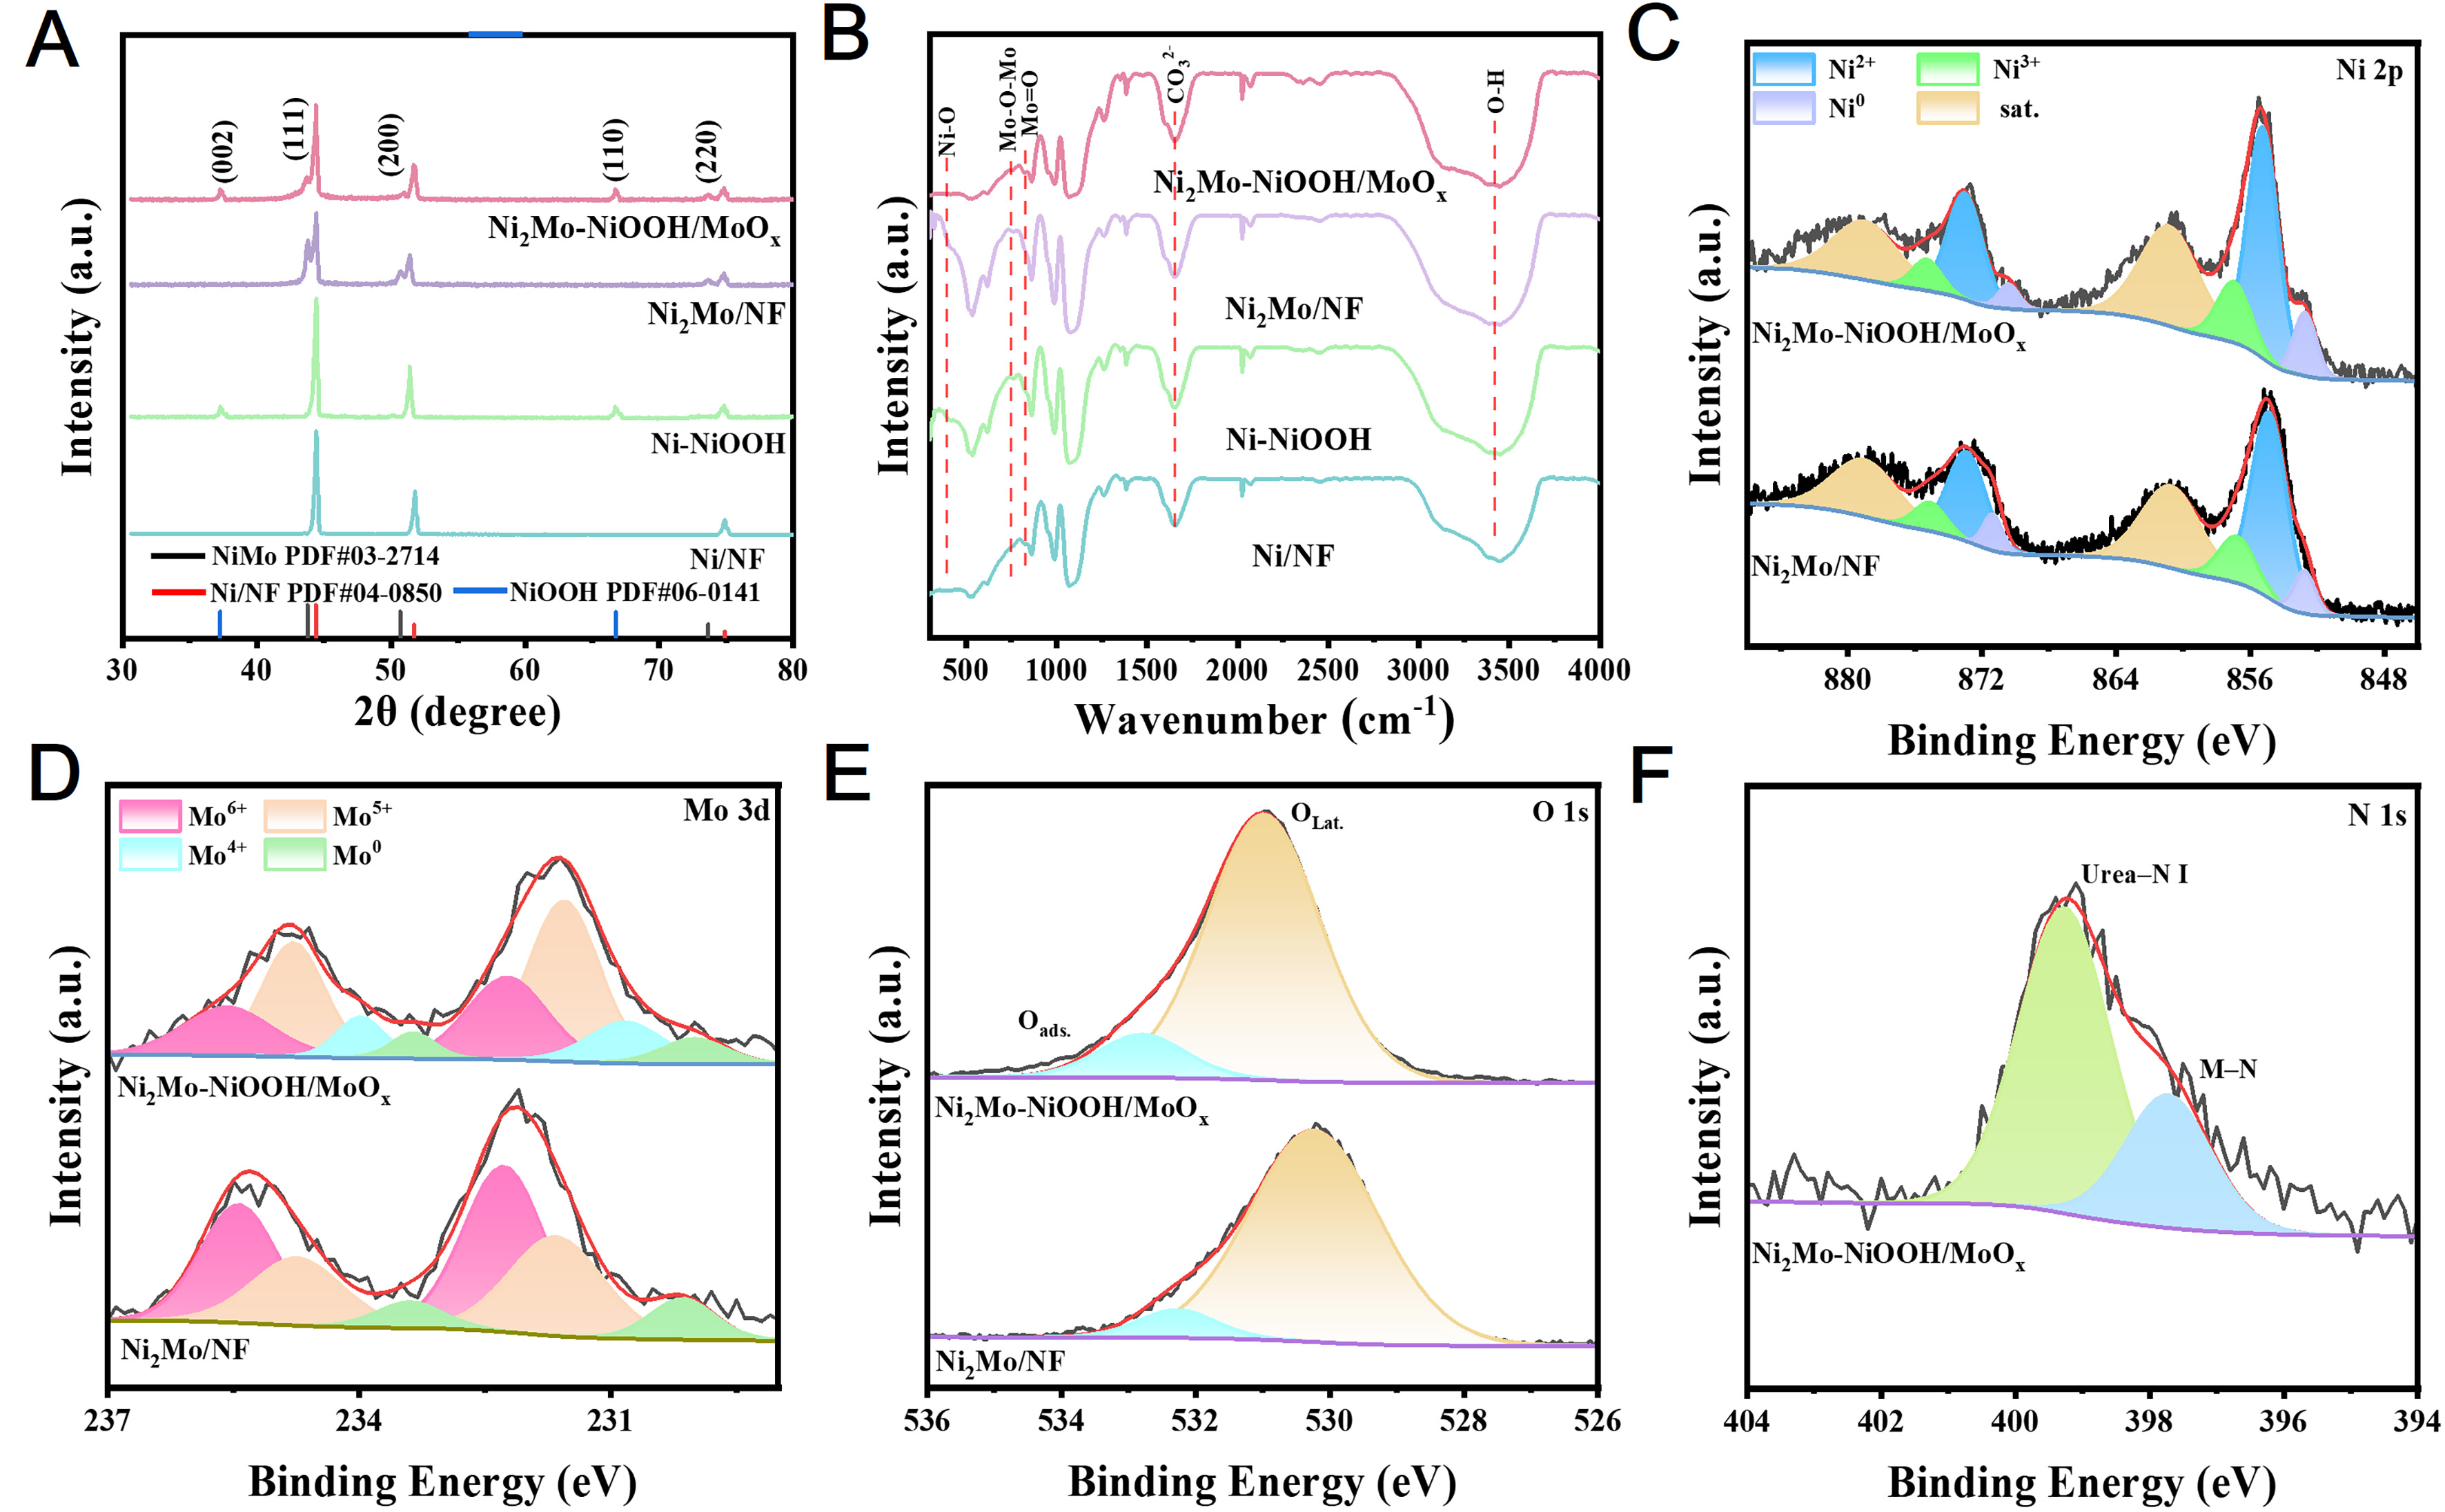
Figure 2. Phase, surface chemical and valence state characterization of catalytic electrodes. (A) XRD patterns of Ni/NF, Ni-NiOOH, Ni2Mo/NF and Ni2Mo-NiOOH/MoOx; (B) FT-IR spectra of the above catalytic electrodes; (C-E) XPS spectra of Ni2Mo/NF and Ni2Mo-NiOOH/MoOx: (C) Ni 2p, (D) Mo 3d, (E) O 1s; (F) N 1s XPS spectrum of Ni2Mo-NiOOH/MoOx. XRD: X-ray diffraction; NF: nickel foam; FT-IR: Fourier transform infrared spectroscopy; XPS: X-ray photoelectron spectroscopy.
X-ray photoelectron spectroscopy (XPS) characterization was carried out to further investigate the electronic structure, elemental valence distribution and electron transfer behavior of the catalysts before and after
In situ Raman analysis during UOR
To reveal the in situ phase evolution and proton-capturing mechanism of the Ni2Mo-NiOOH/MoOx catalytic electrode in the UOR, in situ Raman spectroscopy was adopted to real-time track the surface structure and species evolution of the catalyst during anodic polarization. Figure 3 compares Ni-NiOOH and Ni2Mo-NiOOH/MoOx in terms of spectral features in different wavenumber ranges and three-dimensional in situ Raman profiles under UOR conditions. The region of 440-600 cm-1 is assigned to the bending and stretching vibration signals of Ni-O bonds in NiOOH. For the Ni-NiOOH catalytic electrode [Figure 3A], obvious characteristic Raman peaks of NiOOH can only be observed at a relatively high anodic polarization potential, and the peak intensity increases slowly with rising potential. This indicates that without amorphous MoOx, the in situ reconstruction of Ni sites into the high-activity NiOOH phase exhibits a delayed process with a high phase-transition energy barrier, which is unfavorable for the rapid initiation of UOR. In contrast, Ni2Mo-NiOOH/MoOx presents distinct NiOOH characteristic signals even at a much lower applied potential [Figure 3D], and the peak intensity grows rapidly as the anodic potential increases. This confirms that the introduction of amorphous MoOx effectively reduces the energy barrier for in situ reconstruction of the active phase. Meanwhile, amorphous MoOx induces local charge rearrangement, modulates the electronic structure of Ni active centers, and optimizes the adsorption configuration and adsorption strength of UOR intermediates. Characteristic signals associated with MoOx-related species can be further identified in the high wavenumber range of 980-1,020 cm-1. Combined with the two-dimensional stacked in situ Raman spectra [Supplementary Figure 3], the Ni-NiOOH system shows no obvious characteristic response in this wavenumber region throughout the test [Supplementary Figure 3A]. By comparison, the Ni2Mo-NiOOH/MoOx sample [Supplementary Figure 3B] displays well-resolved characteristic Raman peaks that gradually strengthen with increasing potential. These signals are attributed to the stretching vibrations of terminal Mo=O double bonds and coordinatively unsaturated Mo-O- species in the MoOx phase, which directly proves that the anodic UOR environment can induce the in situ generation of MoOx active components rich in Mo=O/Mo-O- on the catalyst surface. Moreover, the characteristic peaks of carbonate intermediates derived from UOR evolve synchronously within 1,040-1,100 cm-1 [Figure 3C and Figure 3F]. Comparative results demonstrate that Ni2Mo-NiOOH/MoOx possesses a lower onset potential and a more prominent signal response for intermediate peaks. It indicates that Mo=O/Mo-O- sites can rapidly capture H+ released from the dehydrogenation of *-NH2 during urea oxidation, timely alleviate interfacial proton accumulation, markedly lower the energy barrier of the key PCET step, and accelerate the dehydrogenation and intermediate transformation kinetics of UOR. The three-dimensional in situ Raman spectra in Figure 3B and Figure 3E further verify that the NiOOH active phase signals, MoOx characteristic peaks at
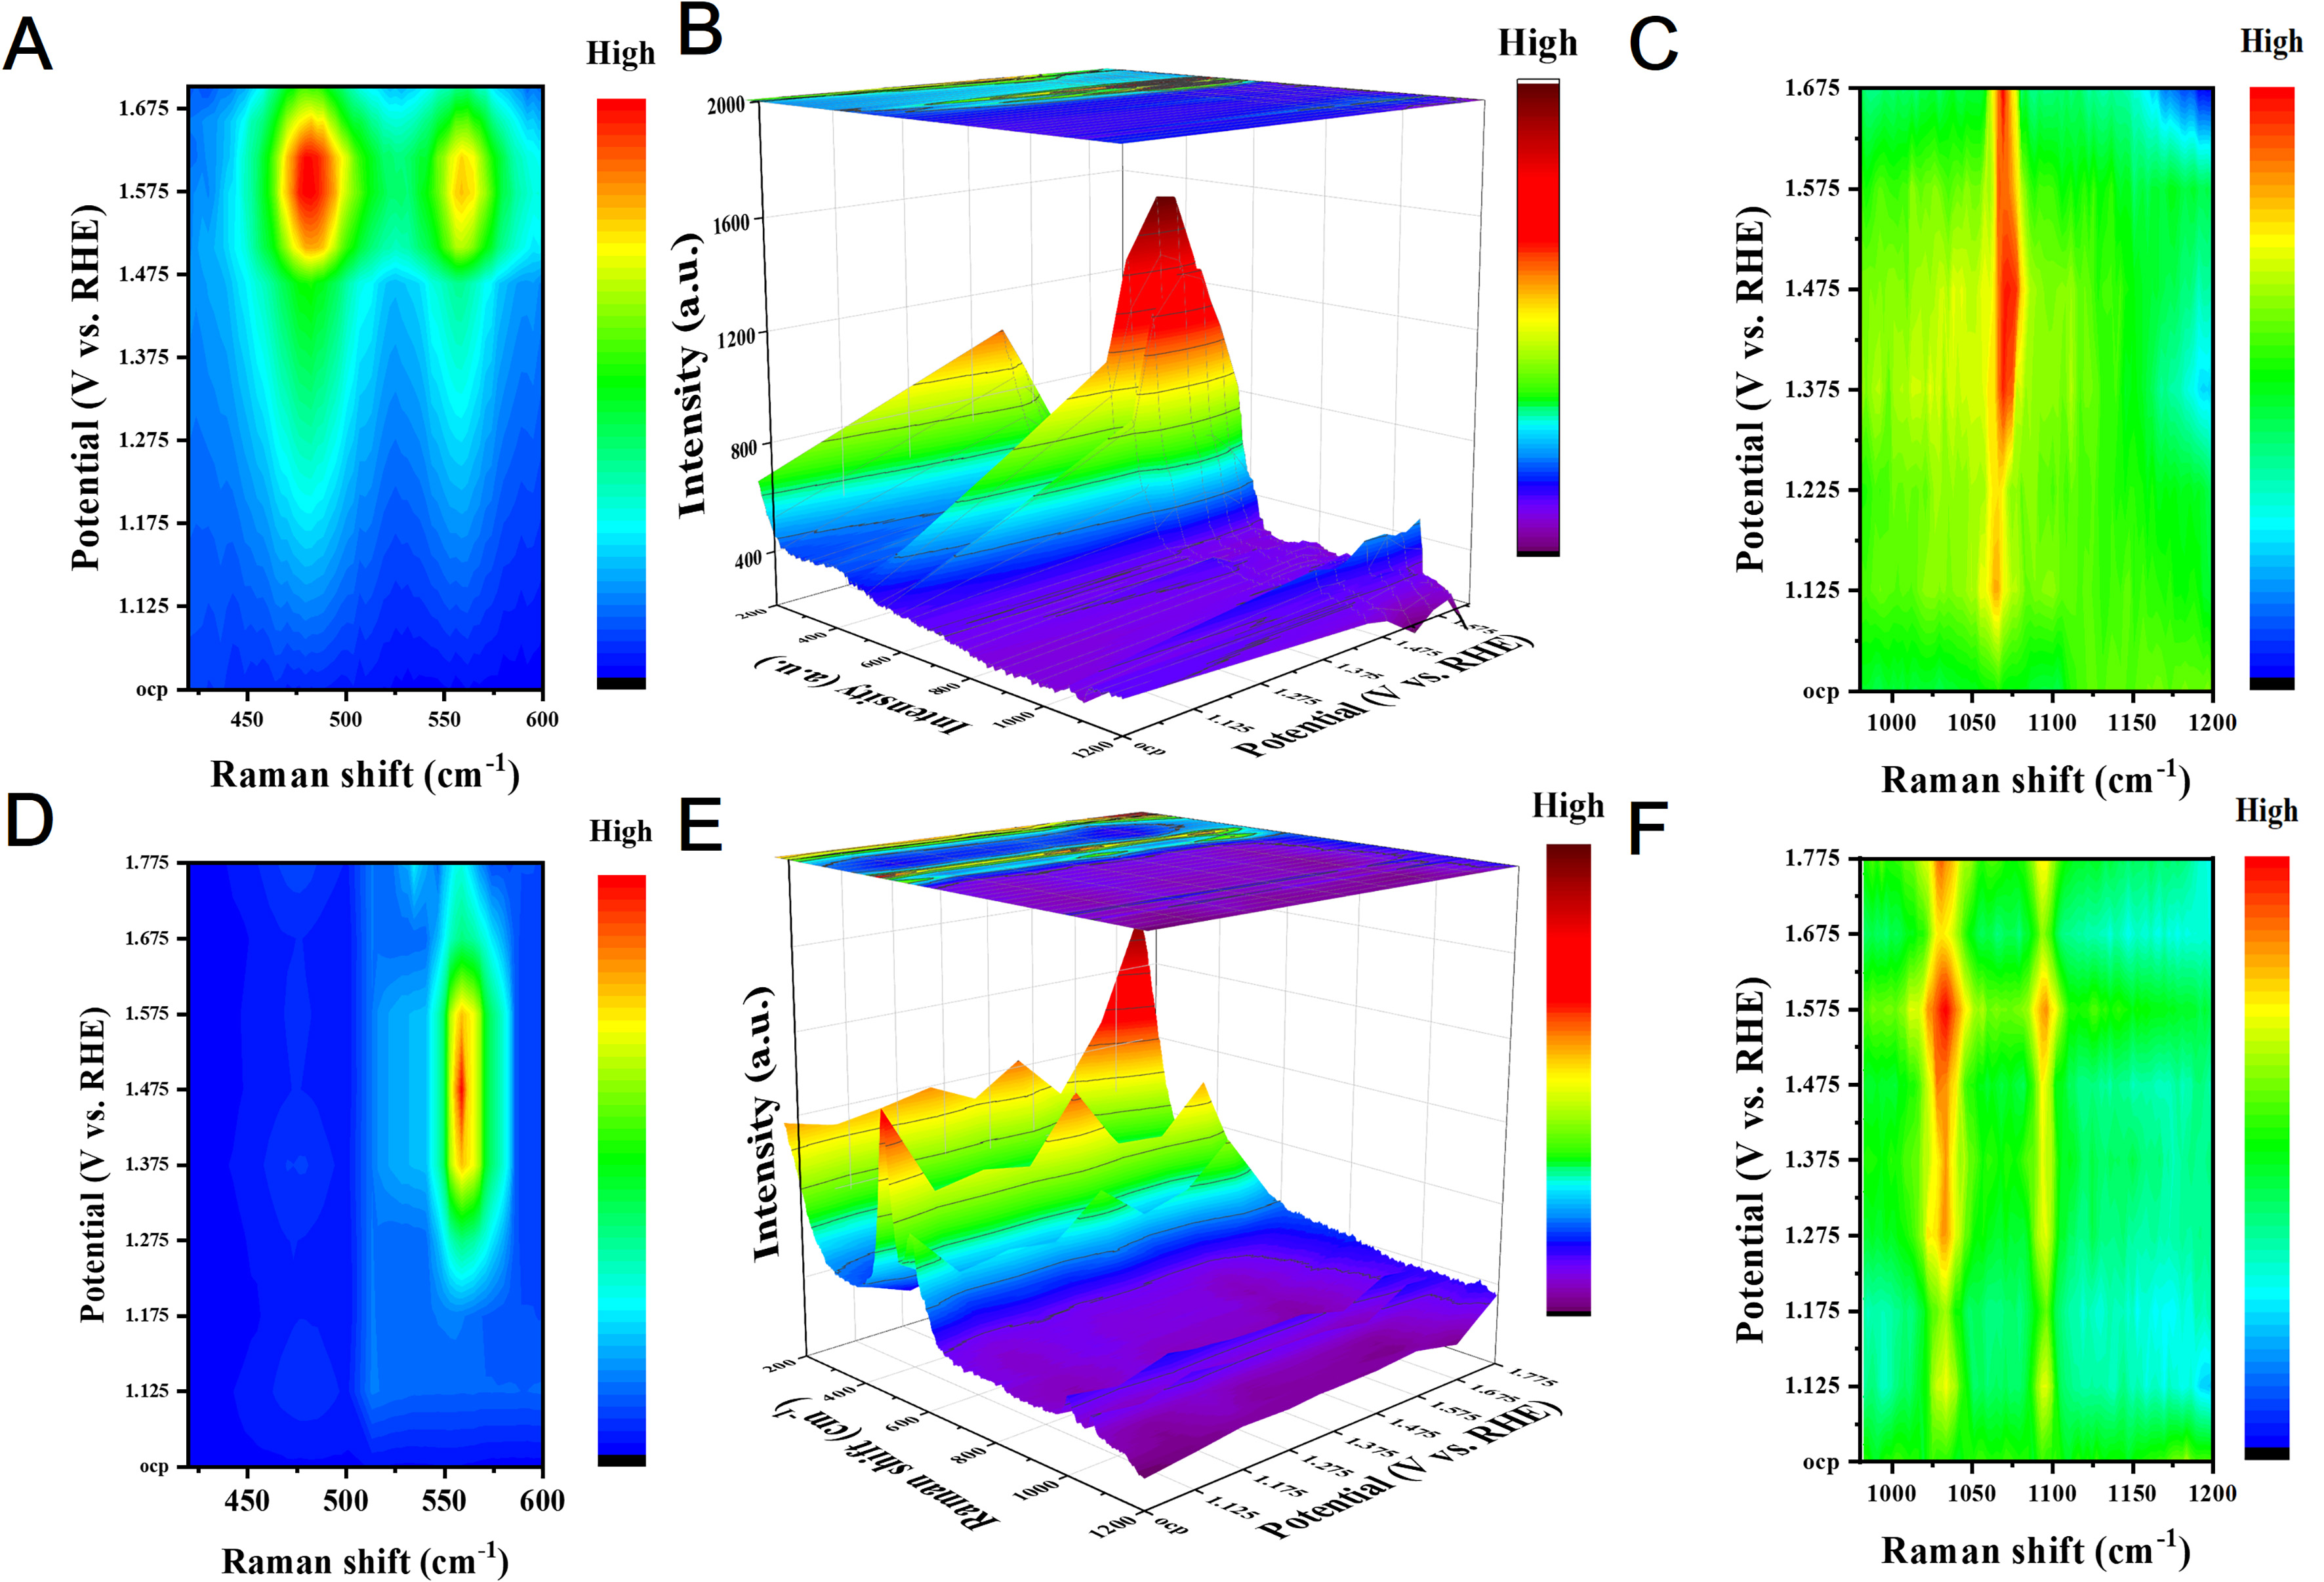
Figure 3. Operando Raman spectra of catalytic electrodes under UOR. (A-C) Operando Raman spectra of Ni-NiOOH at varied anodic potentials: (A) 2D contour plot in the range of 440-600 cm-1; (B) corresponding 3D waterfall plot; (C) 2D contour plot in the range of 980-1,200 cm-1; (D-F) Operando Raman spectra of Ni2Mo-NiOOH/MoOx at varied anodic potentials: (D) 2D contour plot in the range of 440-600 cm-1; (E) corresponding 3D waterfall plot; (F) 2D contour plot in the range of 980-1,200 cm-1. RHE: Reversible hydrogen electrode; UOR: urea oxidation reaction.
In summary, the in situ Raman results confirm that a Ni2Mo-NiOOH/MoOx composite active interface is
Electrochemical performance analysis for UOR
Electrochemical measurements were performed to verify the synergistic effect between the in situ reconstructed Ni2Mo-NiOOH/MoOx catalytic electrode and MoOx proton-capturing sites from the perspectives of catalytic activity, reaction kinetics and reaction selectivity. A standard three-electrode configuration was adopted throughout the electrochemical tests. The UOR performance of the catalysts was evaluated in 1.0 M KOH solution containing 0.33 M urea, while the OER performance was measured in pure 1.0 M KOH electrolyte. Optimization experiments on the Ni/Mo precursor molar ratio [Supplementary Figure 5A] and in situ reconstruction duration of Ni2Mo/NF [Supplementary Figure 5B] were systematically conducted. The results demonstrate that Ni2Mo/NF with a Ni/Mo ratio of 2 exhibits the optimal UOR activity. This can be attributed to the balanced synergy between NiOOH active sites and MoOx proton-capturing sites. An insufficient Mo content leads to the formation of fewer MoOx species and limited proton-capturing capability, whereas excessive Mo incorporation reduces the proportion of NiOOH active sites, resulting in an unfavorable catalytic balance. Moreover, the Ni2Mo-NiOOH/MoOx electrode obtained after 60 min of in situ reconstruction delivers the best UOR performance, providing the basis for subsequent electrochemical studies.
The linear sweep voltammetry (LSV) curves in Figure 4A compare the current density of bare Ni/NF, Ni-NiOOH, Ni2Mo/NF and in situ reconstructed Ni2Mo-NiOOH/MoOx at the same applied potential. It is clearly observed that Ni2Mo-NiOOH/MoOx with MoOx proton-capturing sites induced by anodic reconstruction exhibits remarkably superior UOR performance to other control catalysts. It should be noted that the potential differences presented in the inset of Figure 4A were calculated by subtracting the theoretical equilibrium potential of OER [1.23 V vs. reversible hydrogen electrode (RHE)] from the measured UOR operating potentials. Here, the OER equilibrium potential is employed solely as a common reference benchmark to facilitate a comparative evaluation of the operating potentials required by different catalysts. The as-prepared Ni2Mo-NiOOH/MoOx exhibits outstanding UOR activity, requiring only
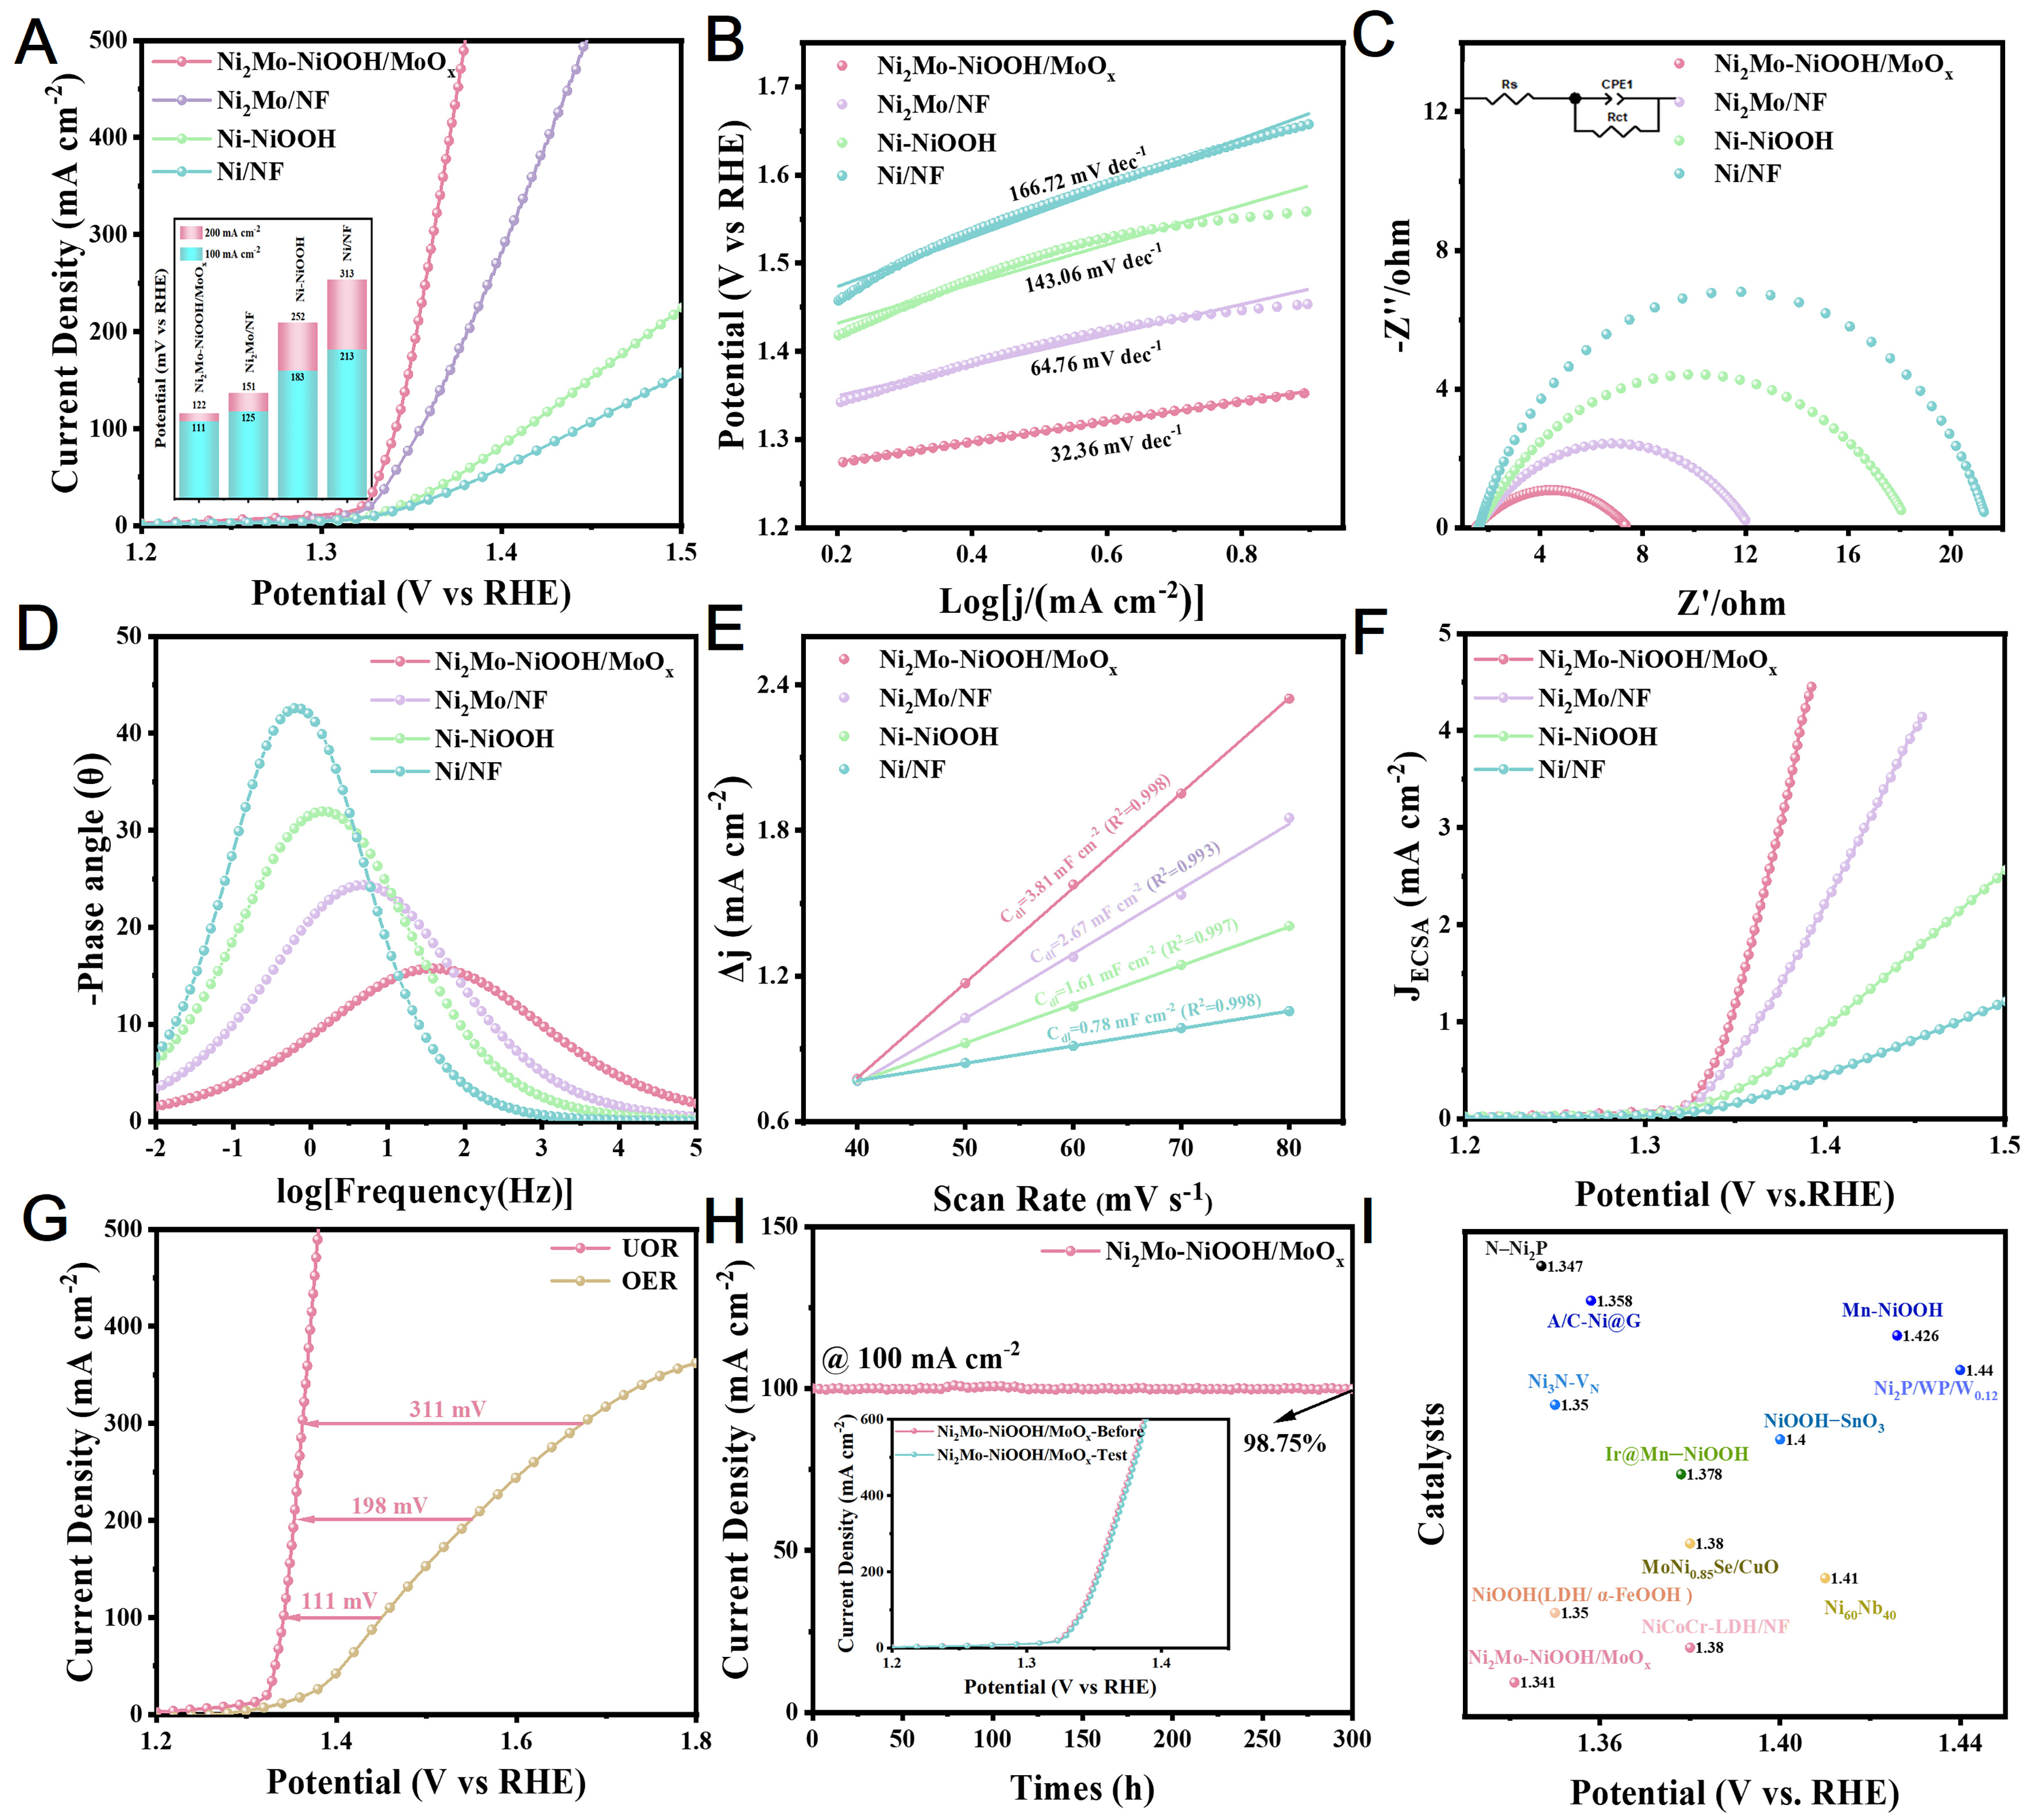
Figure 4. Electrocatalytic UOR performance characterization of various catalysts. (A) LSV curves of Ni2Mo-NiOOH/MoOx, Ni2Mo/NF, Ni-NiOOH and Ni/NF (Inset: Comparison of potential differences relative to the OER equilibrium potential for different catalysts at specific current densities); (B) Corresponding Tafel slope curves (R2 values above 0.95); (C) Nyquist impedance plots and equivalent circuit model; (D) Bode phase plots; (E) Cdl plots (R2 values above 0.99); (F) ECSA-normalized LSV curves; (G) LSV curve comparison between UOR and OER of Ni2Mo-NiOOH/MoOx; (H) 300 h i-t test of Ni2Mo-NiOOH/MoOx at the current density of 100 mA·cm-2, with the LSV curves before and after the stability test shown in the inset; (I) Comparison of the potential of Ni2Mo-NiOOH/MoOx and other UOR catalysts at a current density of 100 mA·cm-2. NF: Nickel foam; LSV: linear sweep voltammetry; UOR: urea oxidation reaction; OER: oxygen evolution reaction; ECSA: electrochemically active surface area; Cdl: double-layer capacitance.
Electrochemical impedance spectroscopy (EIS), phase angle analysis, cyclic voltammetry (CV) and double-layer capacitance (Cdl) measurements were performed to clarify the synergistic enhancement mechanism of the in situ reconstructed Ni2Mo-NiOOH/MoOx composite structure and MoOx proton-capturing sites toward UOR performance from the aspects of charge transport efficiency, active site exposure and intrinsic reaction kinetics.
In terms of charge transport, the Nyquist plots presented in Figure 4C and Supplementary Table 3 show the impedance values of Ni2Mo-NiOOH/MoOx, Ni2Mo/NF, Ni-NiOOH and Ni/NF measured at an open-circuit potential of 0.23 V are 5.81, 10.43, 16.45 and 19.64 Ω, respectively. Among all samples, Ni2Mo-NiOOH/MoOx exhibits the smallest charge transfer resistance (Rct), implying the lowest charge transfer barrier at the electrode/electrolyte interface. This favorable characteristic originates from the local charge rearrangement induced by Mo during in situ reconstruction, which optimizes the electron conduction pathway of the composite architecture and reduces the intrinsic electron transport resistance. Meanwhile, the MoOx proton-capturing sites generated during UOR can rapidly guide interfacial proton migration and avoid interfacial polarization caused by proton accumulation from the dehydrogenation of *-NH2, thereby greatly improving the charge transfer efficiency. Compared with the Nyquist plots under OER conditions [Supplementary Figure 7A], although Ni2Mo-NiOOH/MoOx still delivers a relatively low impedance value, its overall Rct is remarkably higher than that under UOR, and the performance gap with other samples is narrowed. This demonstrates that its low-resistance interfacial property is a specific optimization targeted at the UOR process, rather than a general enhancement of electron conduction. The Bode phase angle plots [Figure 4D and Supplementary Figure 7B] further confirm that the phase angle peak of Ni2Mo-NiOOH/MoOx under UOR appears at a higher frequency region with a broader peak shape. This indicates faster high-frequency charge transfer and a stronger coupling effect between the low-frequency electric double layer and Faradaic reaction, which directly reflects that MoOx proton-capturing sites accelerate interfacial proton delivery and realize efficient coupling of charge and proton transfer. In contrast, the position difference of phase angle peaks among various samples under OER is negligible, failing to embody the superiority of this synergistic effect.
CV curves recorded at various scan rates under UOR conditions [Supplementary Figure 8] show that Ni2Mo-NiOOH/MoOx possesses a much larger enclosed CV area than all control samples, indicative of a higher double-layer capacitance and enhanced electrochemically active surface area (ECSA). Under OER conditions [Supplementary Figure 9], although Ni2Mo-NiOOH/MoOx still exhibits a larger CV area, the overall enhancement degree is far lower than that under UOR, further demonstrating that its active sites are more sufficiently utilized during the UOR process. The Cdl values calculated from CV curves [Figure 4E, Supplementary Figure 10 and Supplementary Table 4] reveal that Ni2Mo-NiOOH/MoOx delivers a markedly higher Cdl than Ni2Mo/NF, Ni-NiOOH and Ni/NF. Given the positive linear correlation between Cdl and ECSA, a larger electrochemically active surface area is confirmed for Ni2Mo-NiOOH/MoOx. This superiority originates from the structural optimization induced by in situ reconstruction. During the transformation of Ni2Mo/NF into the Ni2Mo-NiOOH/MoOx composite structure, abundant active crystal facets and reaction sites are formed on the surface. Meanwhile, the introduction of Mo=O/Mo-O- optimizes the surface hydrophilicity of the electrode, facilitates electrolyte infiltration, and further improves the accessibility of active sites, providing sufficient reaction interfaces for UOR. As displayed in the ECSA-normalized LSV curves [Figure 4F], Ni2Mo-NiOOH/MoOx achieves a higher current density per unit electrochemically active area than other samples. This result indicates that the excellent UOR performance is not merely attributed to the increased number of active sites, but essentially stems from the substantially enhanced intrinsic catalytic activity of individual active sites. The improvement of intrinsic activity strongly verifies the synergistic regulation mechanism between NiOOH active sites and MoOx proton-capturing sites. NiOOH serves as the core oxidative site to drive the adsorption and bond dissociation of urea molecules, while Mo=O/Mo-O- species rapidly capture the released H+ during dehydrogenation, alleviate interfacial proton accumulation, and lower the energy barrier of the PCET step. Moreover, Mo-induced local charge rearrangement modulates the electronic structure of Ni active centers and regulates the adsorption-desorption behavior of UOR intermediates, thereby remarkably boosting the reaction rate of individual active sites.
Figure 4G shows the LSV comparison between UOR and OER, which further reflects the specific performance advantages of the catalyst. At the same current density, Ni2Mo-NiOOH/MoOx exhibits remarkably higher activity in UOR than in OER, accompanied by a much lower overpotential. This superiority stems from the specific adaptation of MoOx proton-capturing sites toward the UOR reaction. The Mo=O/Mo-O- species in MoOx can only accelerate the PCET process by capturing H+ released from the
Figure 4H presents the i-t curve of Ni2Mo-NiOOH/MoOx for continuous steady-state operation over 300 h at 100 mA·cm-2. The current density maintains stable output without obvious attenuation throughout the ultra-long cycling period. Among them, the inset shows the LSV curves before and after the stability test, which almost completely overlap, and no obvious shift of catalytic overpotential is observed, fully demonstrating the excellent electrochemical durability of the catalyst. Such long-term stability benefits from the synergistic superiority of the in situ constructed Ni2Mo-NiOOH/MoOx composite structure. Thermodynamically stable NiOOH serves as the core oxidative active framework, which effectively inhibits the dissolution loss and excessive oxidation of active components under high anodic potential. Meanwhile, the Mo=O/Mo-O- species in MoOx can continuously capture H+ released from *-NH2 dehydrogenation during UOR via reversible proton adsorption-desorption cycles, timely relieve interfacial proton accumulation and weaken local polarization corrosion, thus achieving dynamic self-maintenance of proton-capturing sites. Microscopic characterizations were performed on Ni2Mo-NiOOH/MoOx after 300 h continuous operation. SEM images [Supplementary Figure 12] confirm that the original microscopic morphology is well preserved without particle aggregation, structural collapse or exfoliation of active phases. Elemental mapping verifies the homogeneous distribution of Ni, Mo and O elements on the material surface. Further TEM characterization [Supplementary Figure 13] confirms at the microscopic scale that the catalyst retains an intact lattice structure without phase separation or structural collapse, and Ni, Mo and O elements still maintain a highly uniform distribution, proving the long-term stability of the heterogeneous interface constructed by NiOOH and MoOx. Multi-current step V-t test results [Supplementary Figure 14] show that Ni2Mo-NiOOH/MoOx can rapidly reach potential equilibrium at each current gradient within a wide current density range of 100-400 mA·cm-2, without obvious potential drift or aggravated polarization. Even under harsh working conditions at high current densities, NiOOH active sites and MoOx proton-capturing sites still maintain efficient synergistic coupling. By virtue of Mo-induced charge rearrangement to optimize the electron transport pathway and relying on MoOx to rapidly dissipate enriched protons, the electrode interface always maintains an efficient and stable charge transfer environment under wide-load working conditions. The microscopic characterizations after long-term reaction confirm the intrinsic origin of catalytic stability from the structural perspective. Furthermore, as shown in Figure 4I and Supplementary Table 5, comparison with other modified materials confirms that Ni2Mo-NiOOH/MoOx delivers a particularly remarkable performance enhancement.
Ion dependence analysis
To directly verify the pivotal role of Mo=O/Mo-O- species in the in situ reconstructed MoOx as proton-capturing sites, control experiments were conducted by introducing extra MoO42- and Ni2+ into the electrolyte, and the relevant results are displayed in Figure 5. In the MoO42- addition experiment, the LSV curves of the Ni2Mo-NiOOH/MoOx catalyst only show a negligible response with the increase of MoO42- concentration [Figure 5A], and the curves almost overlap with those obtained in the original electrolyte, presenting an obvious saturation effect. This is because abundant highly active Mo=O/Mo-O- proton-capturing sites have been in situ generated on the electrode surface during anodic reconstruction. These sites are uniformly distributed on the surface of the NiOOH active phase, which can efficiently trap H+ released from *-NH2 dehydrogenation during UOR, alleviate interfacial proton accumulation, and lower the energy barrier of the key PCET step. Therefore, the extra introduced solution-phase MoO42- cannot form more effective proton-capturing sites on the pre-reconstructed electrode surface, resulting in a greatly limited performance improvement space. This directly confirms that the UOR performance of the catalyst mainly originates from the in situ generated MoOx proton-capturing sites rather than free Mo species in the solution. In sharp contrast, the control catalytic electrodes including Ni2Mo/NF, Ni-NiOOH and Ni/NF exhibit an obvious response effect, where the LSV curves shift toward the left and the current density increases remarkably with the rising MoO42- concentration [Figure 5B-D]. This phenomenon arises from the lack of intrinsic MoOx proton-capturing sites on the surface of these catalysts. The dissolved MoO42- in the solution can be in situ converted into Mo=O/Mo-O- contained MoOx species on the electrode surface under anodic potential, providing additional proton-capturing capacity for UOR, accelerating PCET kinetics, and thereby substantially reducing the overpotential and boosting catalytic activity. This result indirectly corroborates the essential function of Mo=O/Mo-O- as proton-capturing sites, and also demonstrates that the MoOx sites derived from in situ reconstruction possess higher stability and efficiency than those formed by Mo species introduced from the bulk solution. In the Ni2+ addition tests [Figure 5E-H], the LSV curves of all catalysts remain nearly unchanged and their performance is barely affected by the variation of Ni ion concentration in the electrolyte. This excludes the interference of solution-phase Ni species on UOR activity and further confirms that the performance modulation is specifically attributed to the efficient proton-capturing behavior of Mo=O/Mo-O- species in MoOx. Such species can synergize with NiOOH active sites to realize the synergistic regulation of electron and proton transfer during UOR, thereby remarkably accelerating the reaction kinetics.
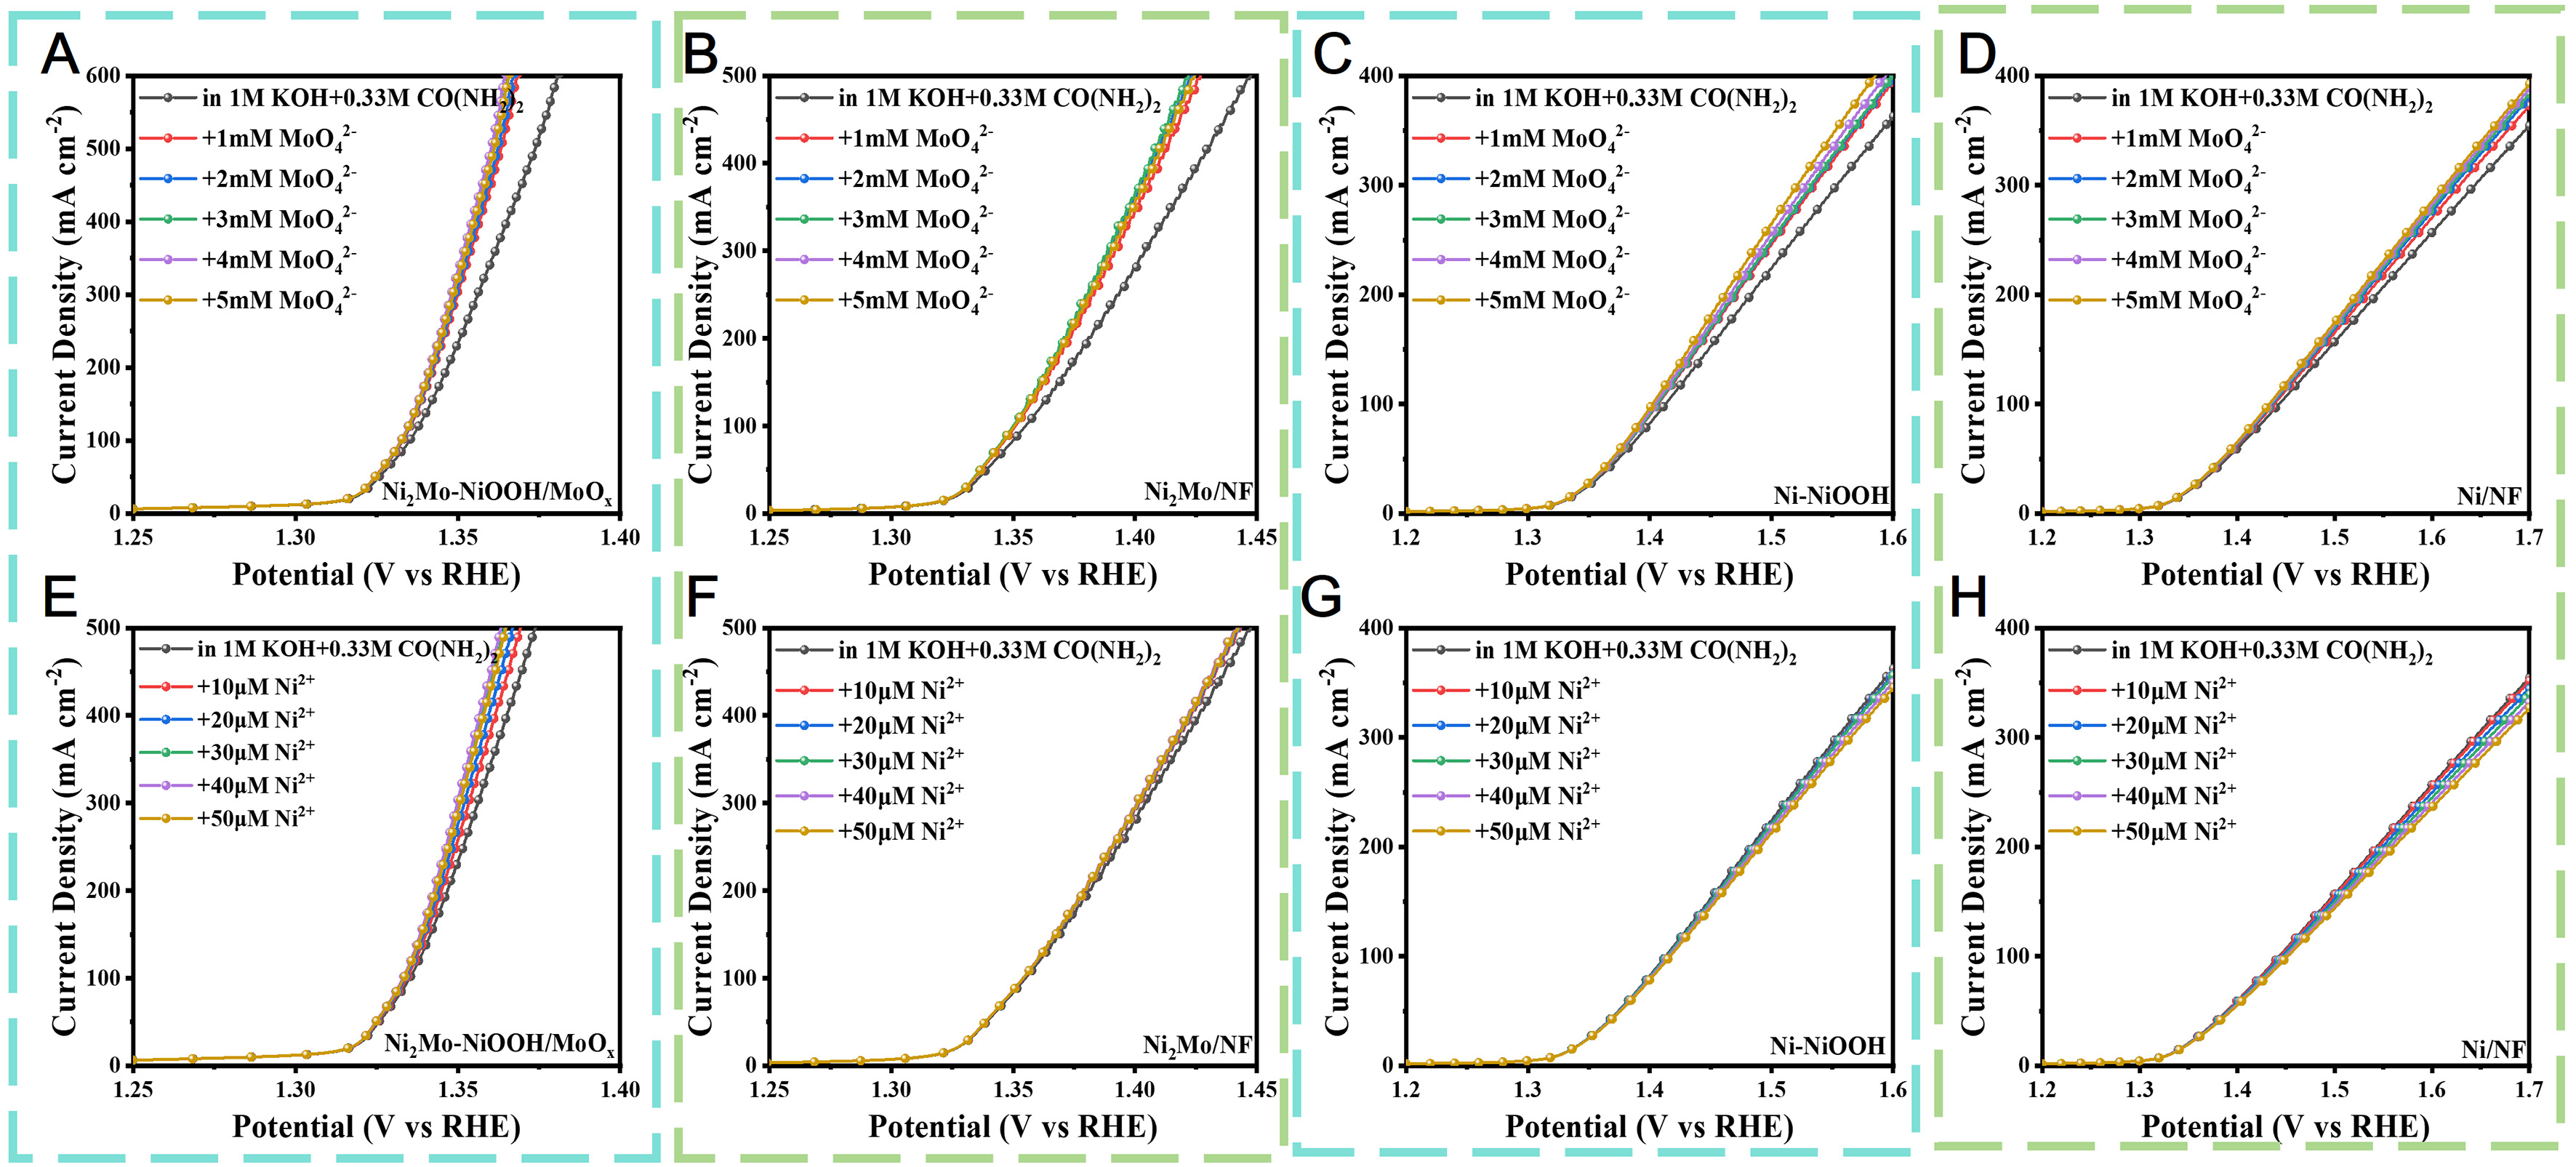
Figure 5. Effects of different concentrations of MoO42- and Ni2+ in the electrolyte on the UOR electrocatalytic performance of various catalytic electrodes. (A-D) LSV curves of Ni2Mo-NiOOH/MoOx, Ni2Mo/NF, Ni-NiOOH and Ni/NF catalytic electrodes with 0-5 mM MoO42- added into the electrolyte containing 1 M KOH and 0.33 M CO(NH2)2; (E-H) LSV curves of the above catalytic electrodes with 0-50 μM Ni2+ added into the electrolyte. RHE: Reversible hydrogen electrode; KOH: potassium hydroxide; LSV: linear sweep voltammetry; UOR: urea oxidation reaction.
Electrolyte concentration-dependent kinetic analysis
To intrinsically reveal the modulation mechanism of Mo=O/Mo-O- proton-capturing sites in MoOx on UOR reaction kinetics, electrolyte concentration-dependent kinetic tests and redox charge capacity characterization were performed[51]. Direct experimental evidence for the synergistic catalytic mechanism was provided from three aspects, including reaction order, interfacial proton transfer efficiency, and active site utilization efficiency.
In the electrolyte concentration-dependent experiments [Figure 6], the RDS of UOR corresponds to the dehydrogenation of urea molecules. The H+ released in this process will accumulate at the electrode interface if not removed timely, inhibiting the forward progression of the reaction and inducing kinetic hysteresis. As shown in Figure 6A and C, the LSV curves of Ni2Mo-NiOOH/MoOx and Ni2Mo/NF were obtained at different urea concentrations. The half-wave current density (j1/2) exhibits a significant linear positive correlation with urea concentration, yielding a higher slope of 616.64 for Ni2Mo-NiOOH/MoOx [Figure 6B], which is much higher than the value of 280.48 for Ni2Mo/NF [Figure 6D]. The stronger concentration dependence indicates that Ni2Mo-NiOOH/MoOx presents a more sensitive response to urea concentration and a remarkably enhanced urea conversion efficiency. The fundamental origin lies in the fact that the in situ generated Mo=O/Mo-O- species can rapidly trap H+ released during dehydrogenation, effectively alleviate interfacial proton accumulation, and drive the forward progression of the PCET process. Accordingly, more urea molecules can be efficiently converted with the increase of urea concentration, and the increment of reaction rate is far larger than that of the precursor catalyst without efficient proton-capturing sites. Kinetic tests targeting KOH concentration further confirm the superiority of MoOx proton-capturing sites. OH- in the solution acts as both the reactant of UOR and the final proton acceptor. As shown in Figure 6E and G, the LSV curves of Ni2Mo-NiOOH/MoOx and Ni2Mo/NF were obtained at different KOH concentrations. The slope calculated from the Log[j] vs. Log[KOH] fitting for Ni2Mo-NiOOH/MoOx is 0.546 [Figure 6F], slightly lower than 0.581 for Ni2Mo/NF [Figure 6H], indicating that this catalyst has a weaker dependence on the OH- concentration in the electrolyte. This difference originates from the reversible proton capture cycle of Mo=O/Mo-O-. Mo=O can in situ capture H+ to form Mo-OH, and subsequently regenerate Mo=O by rapidly reacting with OH-, thus constructing a closed loop of “capture-transfer-regeneration”. Efficient proton conduction can be maintained even at low OH- concentrations. In contrast, Ni2Mo/NF lacks such surface proton-capturing sites and has to completely depend on OH- in the solution to accept H+, rendering it more sensitive to the variation of OH- concentration and a higher reaction order. Redox charge capacity characterization [Supplementary Figure 15] further verifies the Mo-induced synergistic enhancement effect. The integrated charge (Q) calculated from the LSV curve of Ni2Mo-NiOOH/MoOx reaches up to
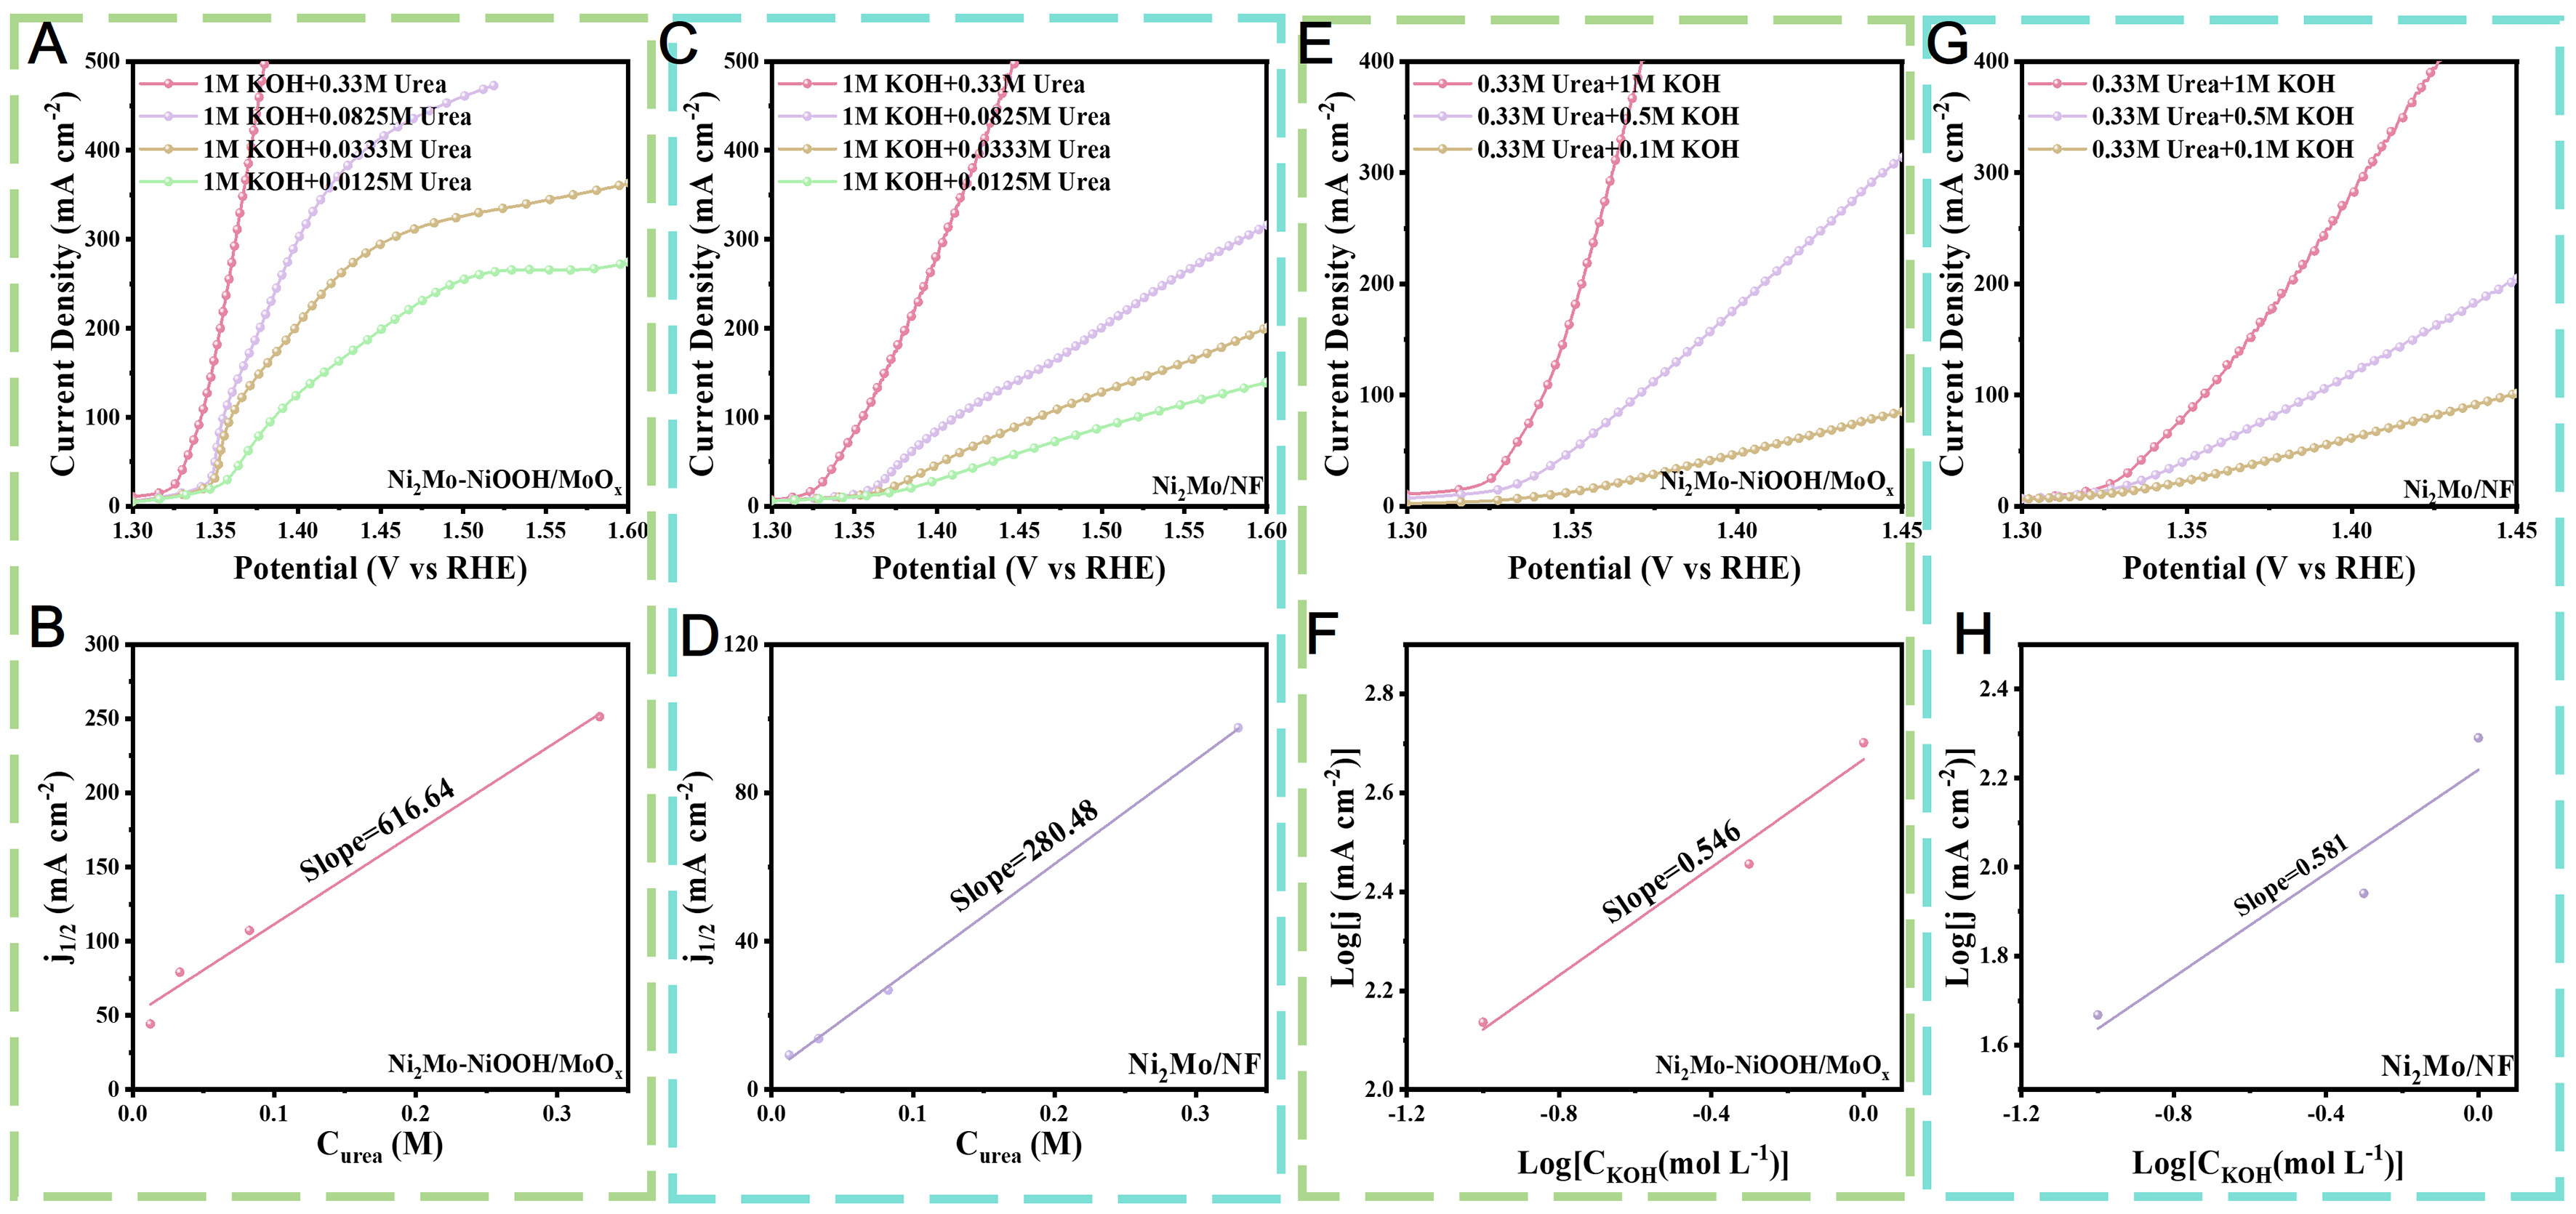
Figure 6. Concentration-dependent kinetic analysis of Ni2Mo-NiOOH/MoOx and Ni2Mo/NF catalysts. (A and C) LSV curves of Ni2Mo-NiOOH/MoOx and Ni2Mo/NF at different urea concentrations (0.0125-0.33 M); (B and D) Linear fitting relationships between j1/2 and urea concentration for Ni2Mo-NiOOH/MoOx and Ni2Mo/NF, respectively, reflecting the concentration dependence toward urea oxidation; (E and G) LSV curves of Ni2Mo-NiOOH/MoOx and Ni2Mo/NF at different KOH concentrations (0.1-1 M); (F and H) Log(j)-Log[KOH] fitting curves for Ni2Mo-NiOOH/MoOx and Ni2Mo/NF, respectively, used to determine the apparent reaction order with respect to OH- concentration. RHE: Reversible hydrogen electrode; NF: nickel foam; KOH: potassium hydroxide; LSV: linear sweep voltammetry; j1/2: half-wave current density.
In summary, electrolyte concentration-dependent kinetic measurements and redox charge capacity characterization validate the core role of MoOx proton-capturing sites in Ni2Mo-NiOOH/MoOx. By rapidly trapping H+ released in the dehydrogenation step of UOR, these sites alleviate interfacial proton accumulation and lower the energy barrier of the PCET step. Meanwhile, they optimize the electronic structure and utilization efficiency of Ni active sites, realizing the synergistic regulation of electron and proton transfer. This work provides solid experimental support for its superior UOR kinetic performance.
In situ electrochemical analysis
To reveal the dynamic evolutionary kinetics of interfacial charge transfer and proton transport at the electrode interface during UOR, in situ electrochemical EIS measurements were conducted at different potentials. Three-dimensional waterfall plots of Nyquist impedance spectra and Bode phase diagrams were utilized to real-time track the potential-dependent evolution of interfacial reaction behavior, providing in situ kinetic evidence for the regulatory effect of MoOx proton-capturing sites.
As presented in Figure 7A, the Rct of Ni2Mo-NiOOH/MoOx decreases remarkably with the increase of potential and shows an obvious decline even in the low-potential region, reaching the minimum value among all catalysts at 0.65 V. This phenomenon directly confirms the crucial role of in situ generated MoOx species. During UOR, the dehydrogenation of urea-derived *-NH2 intermediates releases a large quantity of H+. If these protons cannot be removed promptly, they will accumulate at the electrode interface, induce interfacial polarization, and substantially increase the Rct. In contrast, the Mo=O/Mo-O- species on the surface of Ni2Mo-NiOOH/MoOx can rapidly capture H+ released from the dehydrogenation process, construct interfacial proton transfer pathways, and effectively alleviate interfacial proton accumulation and polarization effects. Consequently, the Rct is greatly reduced, establishing a low-resistance interfacial environment for the PCET process. In comparison, the Rct of Ni2Mo/NF [Figure 7B] varies slowly in the low-potential range and remains distinctly higher than that of Ni2Mo-NiOOH/MoOx, indicating that stable and efficient MoOx proton-capturing sites are not formed and the capacity of interfacial proton conduction is insufficient. Furthermore, the Rct variation of the Ni-NiOOH catalyst [Figure 7C] exhibits a more delayed trend, with only a limited decline at high potentials. This further verifies that the lack of proton-capturing sites leads to blocked charge transfer caused by interfacial proton accumulation and severe retardation of reaction kinetics. Bode phase diagrams [Figure 7D-F] further clarify the kinetic characteristics of interfacial reactions. The phase angle peak of Ni2Mo-NiOOH/MoOx shifts toward the high-frequency direction and continuously increases in intensity as the potential rises, implying a greatly shortened time constant of interfacial reactions and a markedly enhanced Faradaic reaction rate. The high-frequency shift of the phase angle peak directly reflects the acceleration effect of MoOx proton-capturing sites on the PCET process. The Mo=O/Mo-O- mediated rapid proton transfer effectively reduces the coupling resistance between charge transfer and proton transport, thereby lowering the kinetic time constant of UOR Faradaic reactions. Meanwhile, the continuous enhancement of peak intensity suggests that the Faradaic contribution of UOR increases gradually with elevated potential, enabling the catalyst to efficiently drive the reaction even at low applied potentials. On the contrary, the amplitude of the high-frequency shift of the phase angle peak for Ni2Mo/NF is much smaller than that of Ni2Mo-NiOOH/MoOx, with a broadened peak shape and low intensity at low potentials, which reflects its limited interfacial proton transport efficiency and inadequate acceleration of the PCET process. The phase angle peak of Ni-NiOOH barely shifts within the entire potential range and maintains a consistently low intensity, further confirming that interfacial proton accumulation seriously retards reaction kinetics without the participation of proton-capturing sites.
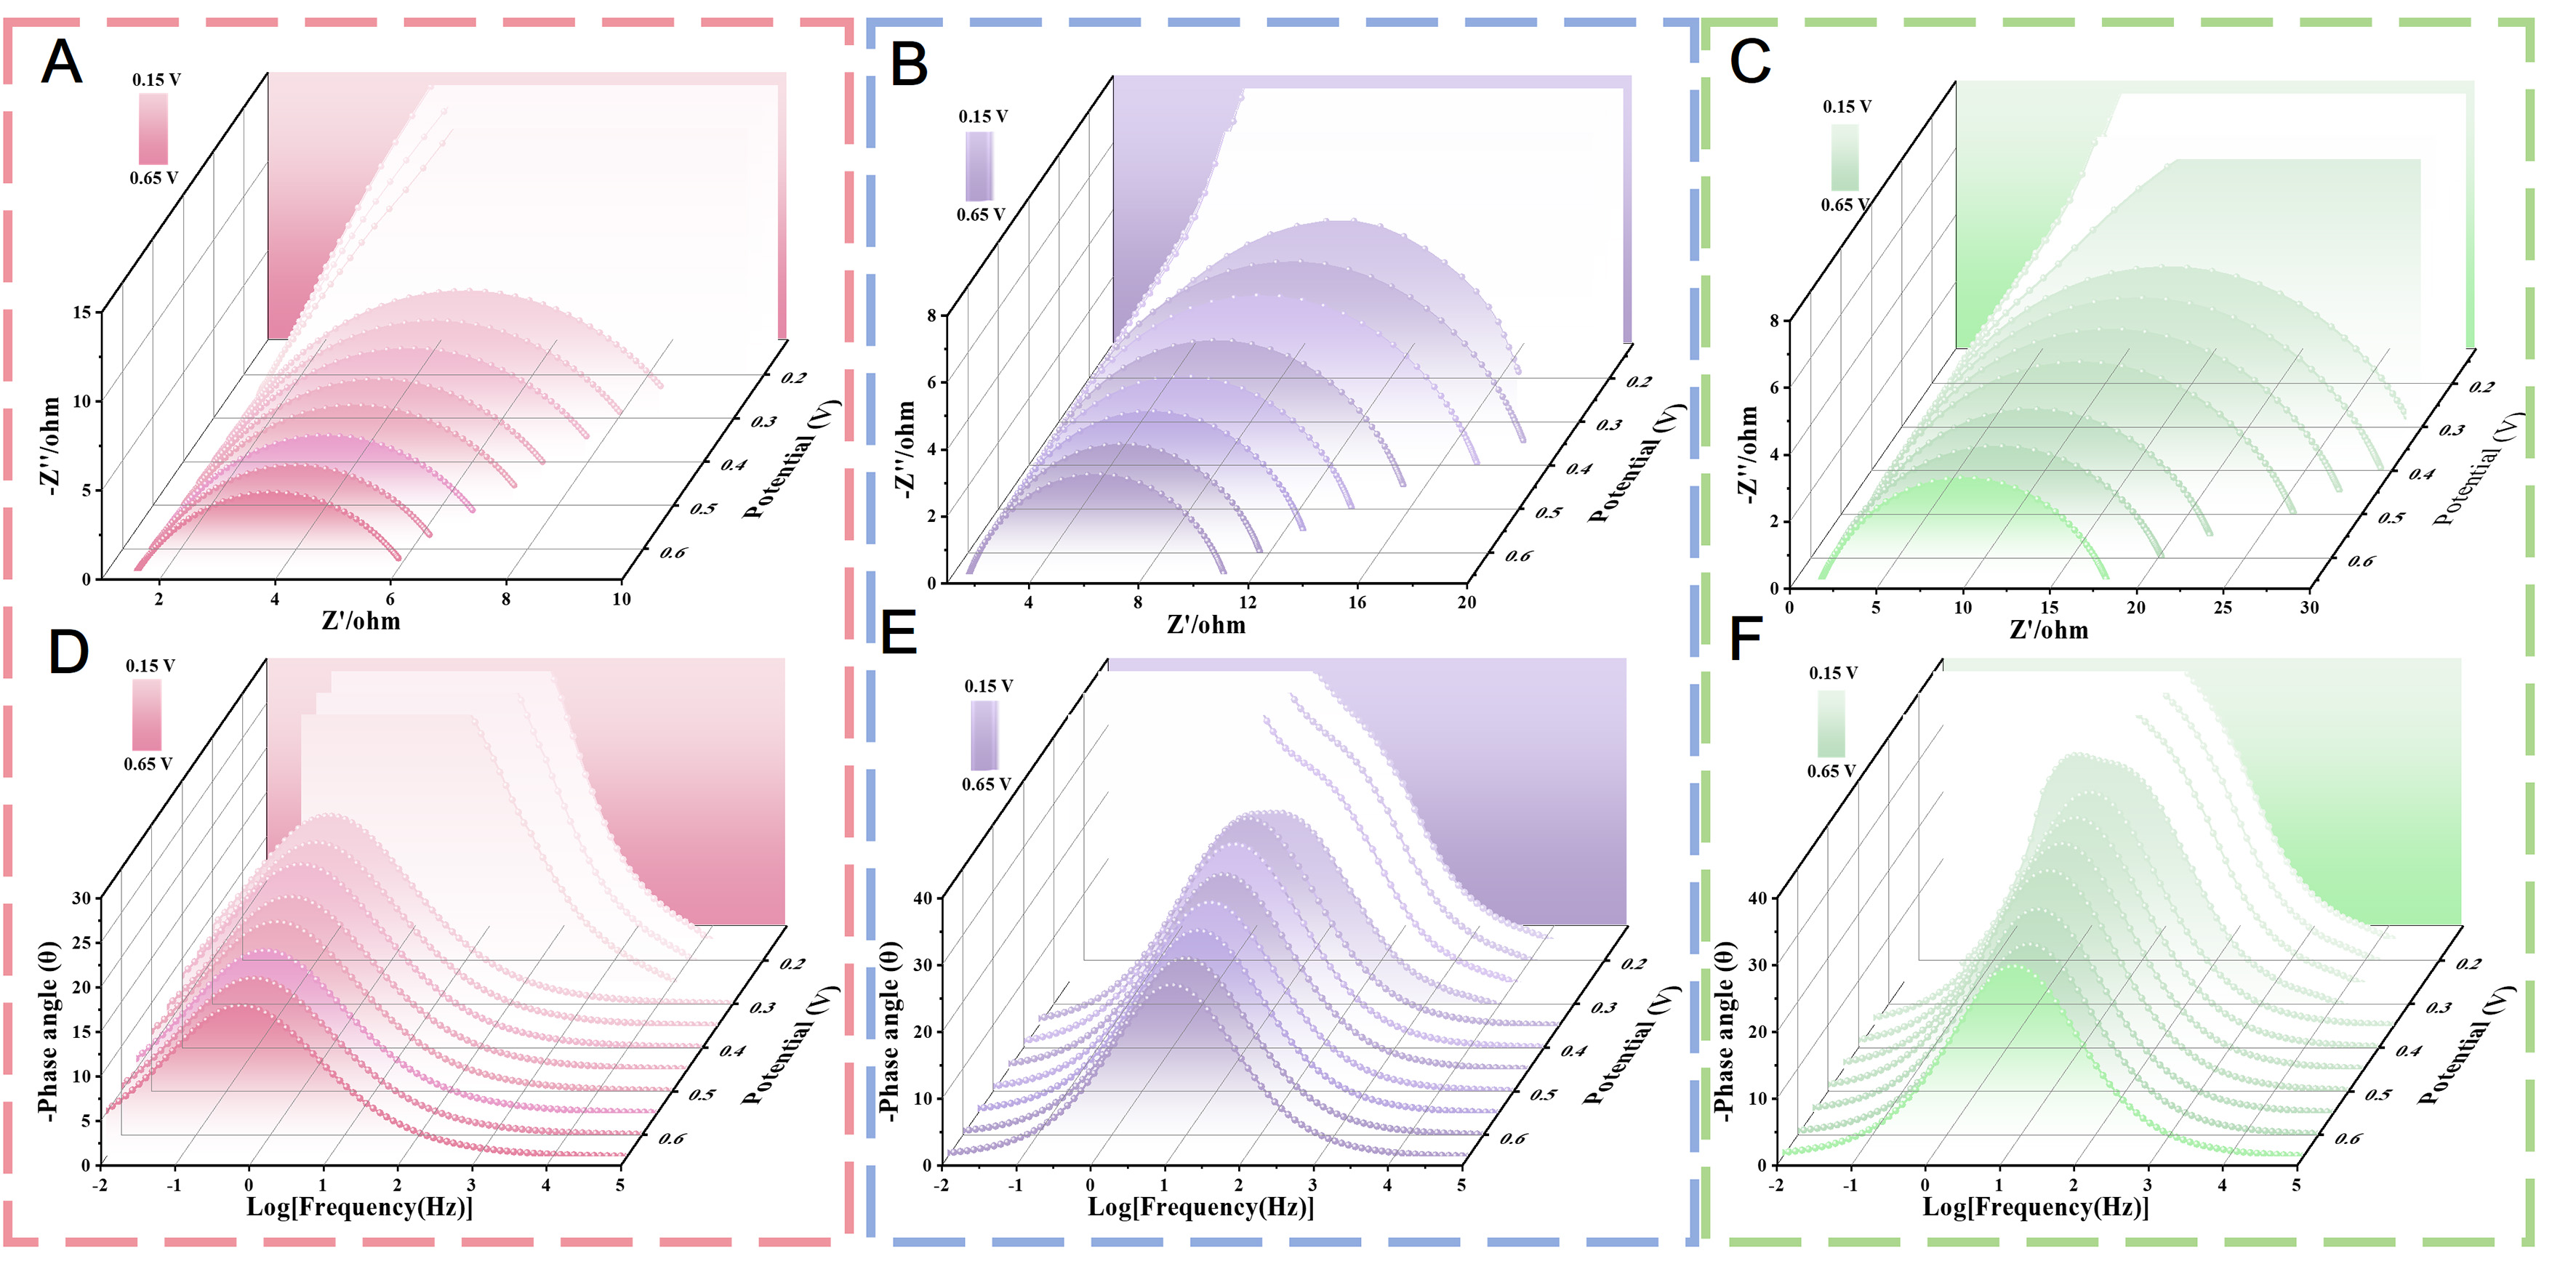
Figure 7. Operando EIS characterization of various catalytic electrodes (potential range: 0.15-0.65 V vs. SCE). (A-C) 3D waterfall plots of Nyquist impedance spectra; (D-F) 3D waterfall plots of Bode phase diagrams; (A and D) Ni2Mo-NiOOH/MoOx; (B and E) Ni2Mo/NF; (C and F) Ni-NiOOH. EIS: Electrochemical impedance spectroscopy; SCE: saturated calomel electrode; NF: nickel foam.
In summary, the dynamic characterization results of in situ EIS three-dimensional waterfall plots fully reveal the pivotal regulatory role of MoOx proton-capturing sites in Ni2Mo-NiOOH/MoOx. By rapidly capturing H+ released during UOR dehydrogenation and constructing interfacial proton transfer pathways, these sites effectively mitigate interfacial polarization and proton accumulation, and synergistically optimize the kinetics of charge transfer and proton transport together with NiOOH active sites.
Purification performance of high-concentration urea wastewater
To evaluate the practical application potential of the Ni2Mo-NiOOH/MoOx electrocatalyst in urea-containing wastewater treatment and coupled urea oxidation systems, the diacetyl monoxime-antipyrine method (Chinese national standard method) was employed to measure the urea degradation efficiency[52]. The urea degradation performance of various catalysts was systematically investigated through ultraviolet–visible spectroscopy (UV-Vis) absorption spectroscopy, solution color variation and real-time monitoring of urea concentration [Figure 8].
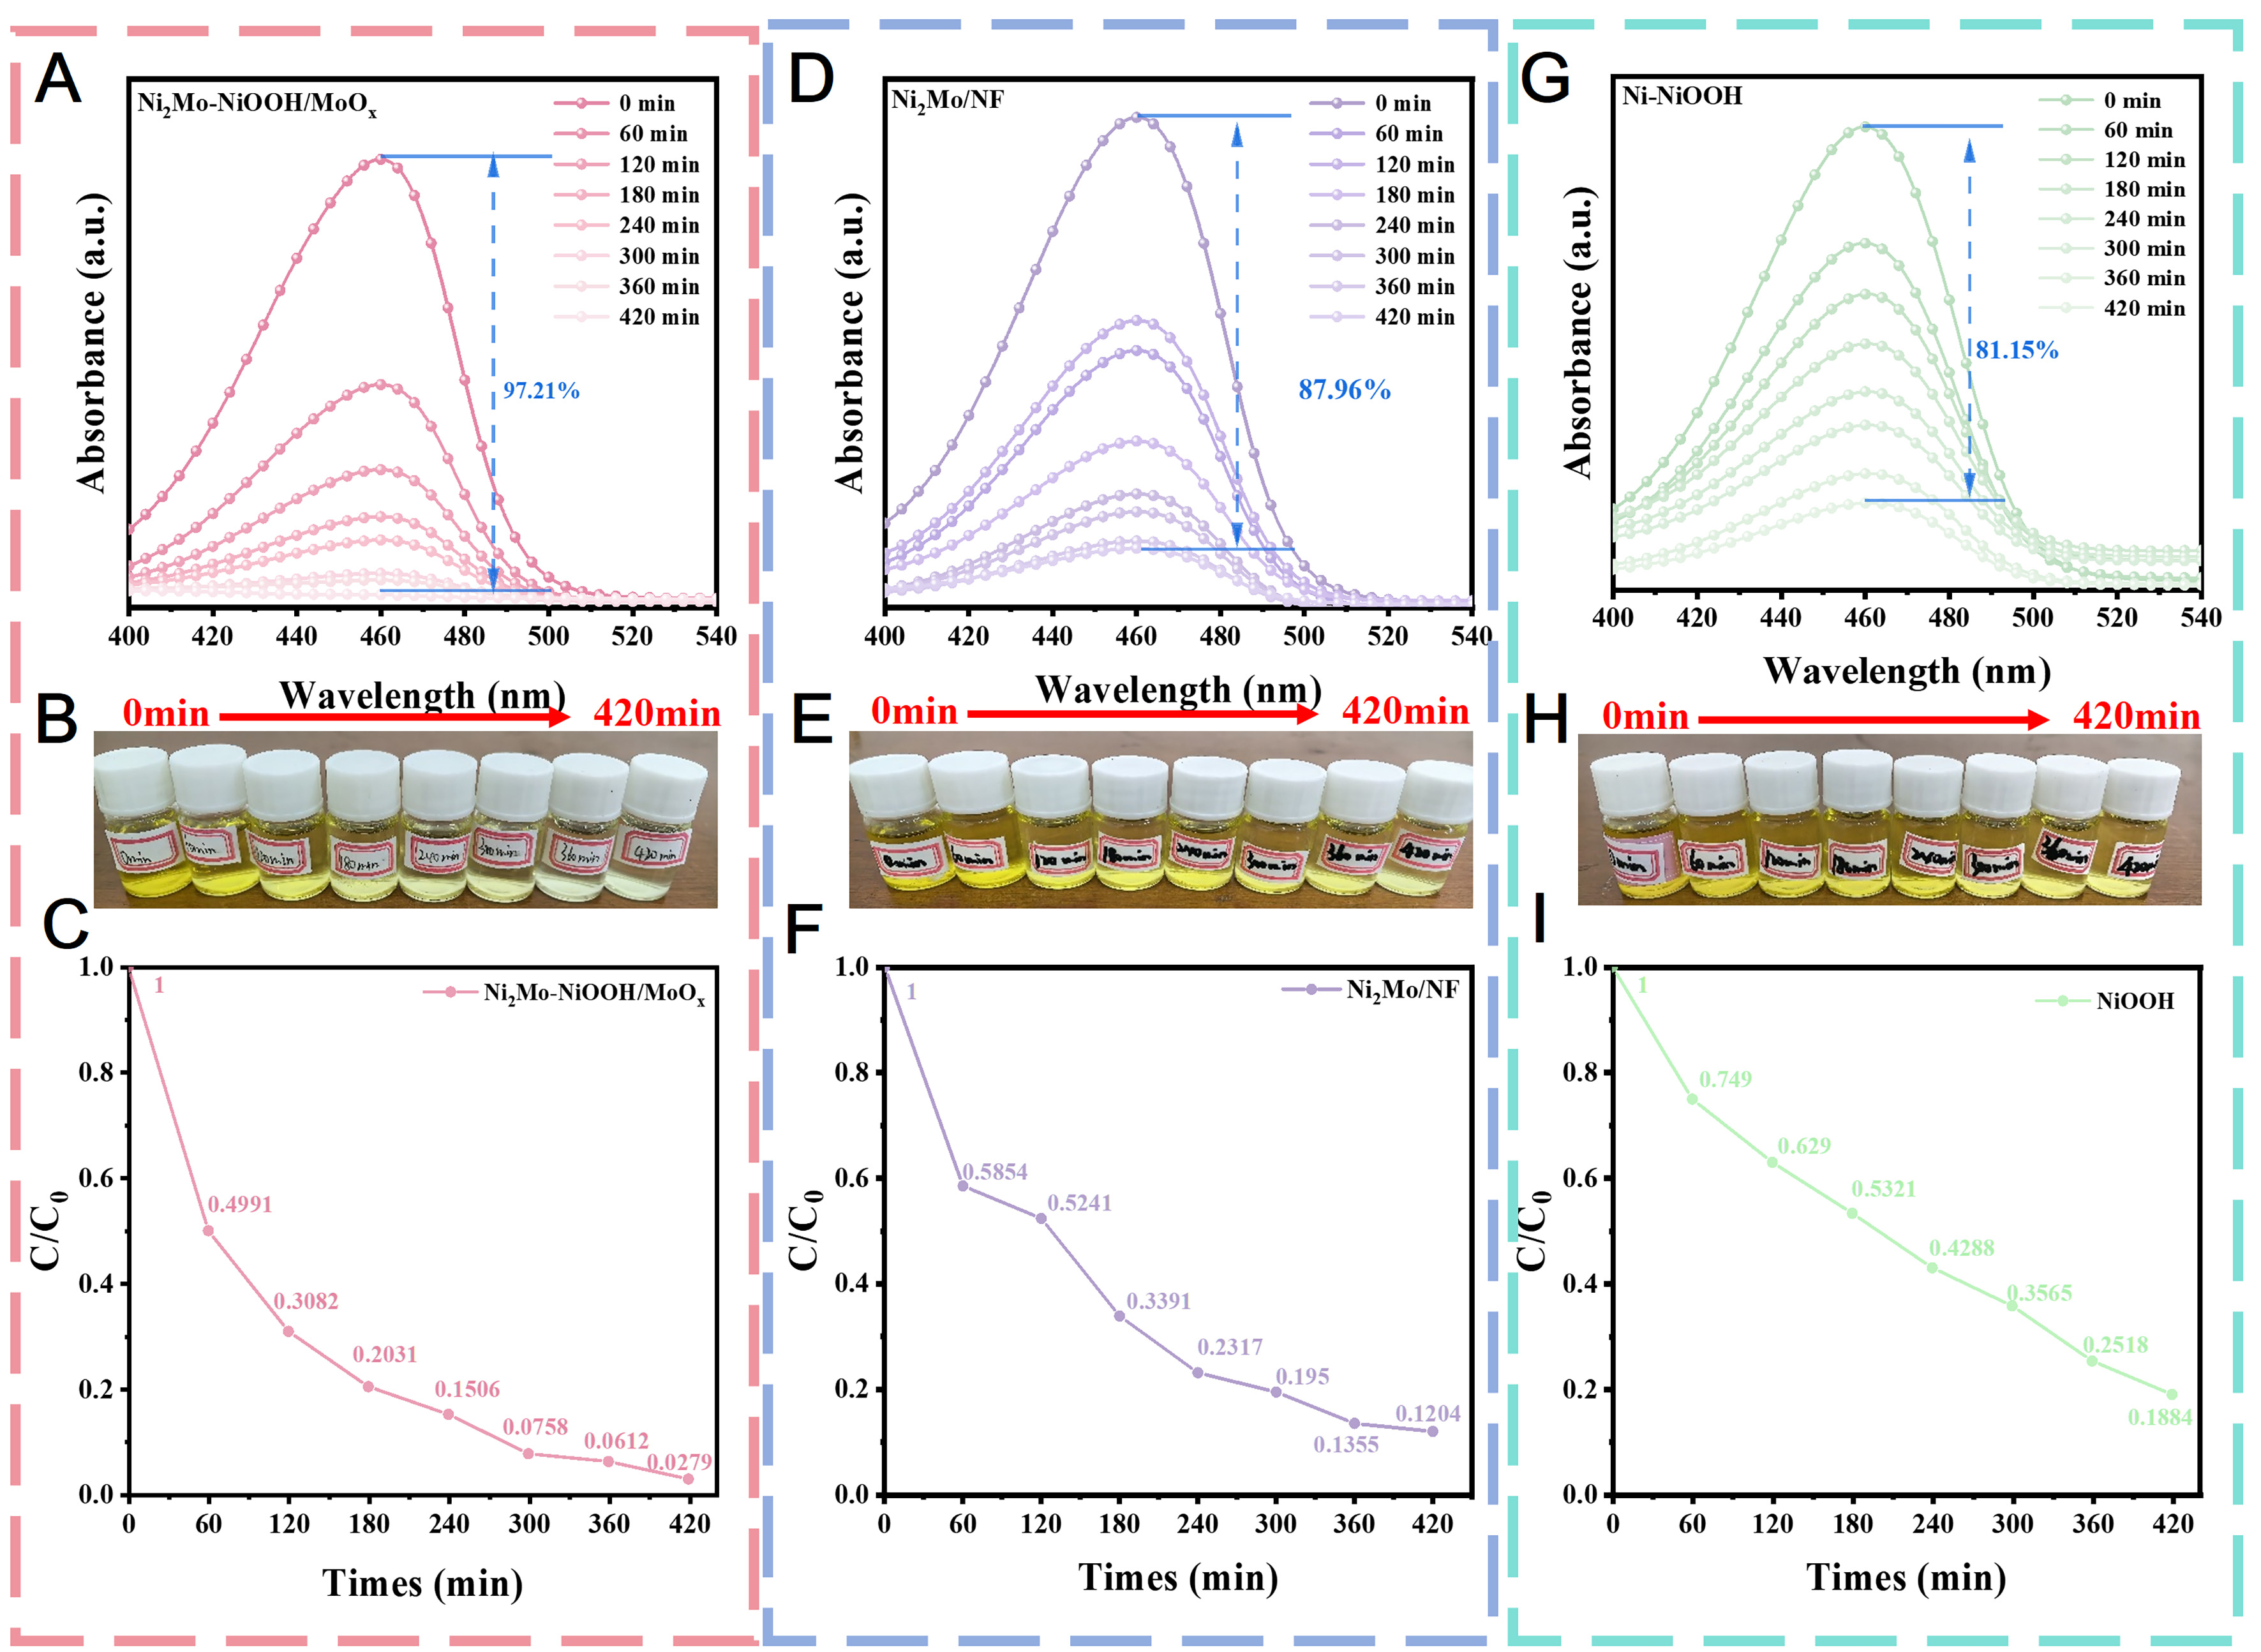
Figure 8. Urea degradation performance characterization of different catalytic electrodes. (A, D, G) UV-Vis absorption spectra at different reaction times; (B, E, H) Photographs of solution color evolution during the reaction; (C, F, I) Variation curves of relative urea concentration C/C0 as a function of reaction time; where (A)-(C), (D)-(F), and (G)-(I) correspond to the test results of Ni2Mo-NiOOH/MoOx, Ni2Mo/NF, and Ni-NiOOH catalysts, respectively. NF: Nickel foam.
As shown in Figure 8A, the characteristic absorption peak corresponding to urea degradation in the Ni2Mo-NiOOH/MoOx system decays rapidly with prolonged reaction time. After 420 min, the absorption intensity drops to only 2.79% of the initial value, corresponding to an ultrahigh urea removal efficiency of 97.21%. This value is remarkably superior to those of Ni2Mo/NF (87.96%) [Figure 8D] and Ni-NiOOH (81.16%) [Figure 8G]. The solution color changes intuitively confirm such performance differences [Figure 8B, E and H]. The yellow color of the solution in the Ni2Mo-NiOOH/MoOx system fades rapidly as the reaction proceeds, while obvious color residue can still be observed in the control catalyst systems, directly reflecting Ni2Mo-NiOOH/MoOx system outstanding urea degradation capability. The time-dependent relative urea concentration curves [Figure 8C, F and I] further quantitatively reveal the kinetic differences. Ni2Mo-NiOOH/MoOx exhibits a much faster decline rate of urea concentration, with the C/C0 value decreasing to 2.79% within 420 min. By contrast, the C/C0 values of Ni2Mo/NF and Ni-NiOOH remain as high as 12.04% and 18.84%, respectively, demonstrating the optimal urea conversion efficiency and reaction rate of Ni2Mo-NiOOH/MoOx. Such prominent superiority fundamentally originates from the synergistic regulation between NiOOH active sites and MoOx proton-capturing sites. As the oxidative active center, NiOOH efficiently drives the adsorption and bond dissociation of urea molecules. Meanwhile, the in situ generated Mo=O/Mo-O- species can rapidly trap H+ released during the *-NH2 dehydrogenation step, effectively alleviate interfacial proton accumulation, markedly lower the energy barrier of the key PCET step, and accelerate UOR kinetics, thus achieving efficient urea degradation under practical working conditions. In comparison, Ni2Mo/NF fails to form stable and efficient MoOx proton-capturing sites, resulting in insufficient interfacial proton dissipation and restricted reaction kinetics. Ni-NiOOH without MoOx proton-capturing sites lacks the proton capture mechanism; the accumulated interfacial protons retard the UOR process and greatly reduce the urea degradation efficiency.
From the perspective of practical application, Ni2Mo-NiOOH/MoOx achieves nearly complete urea removal within 420 min, showing great application potential in urea-containing wastewater treatment, direct urea fuel cells, and urea-assisted hydrogen production coupling systems. This work provides a feasible practical strategy for developing low-cost and high-activity electrocatalysts toward urea oxidation, and further verifies the reliability and superiority of this synergistic catalytic strategy under practical operating conditions.
Figure 9 illustrates the catalytic electrode with dynamically reconstructed Ni2Mo-NiOOH/MoOx, which possesses a heterogeneous interface co-constructed by NiOOH active phases and MoOx species. The NiOOH phase functions as the core oxidative active center, providing reactive sites for the adsorption and stepwise dehydrogenation of urea molecules and driving the forward progression of UOR. Meanwhile, the synchronously generated MoOx is rich in Mo=O/Mo-O- species, which can act as efficient proton-capturing centers. These sites rapidly trap protons released from the *-NH2 dehydrogenation step, effectively alleviate the polarization effect caused by interfacial proton accumulation, and markedly lower the reaction energy barrier of the key PCET step. Consequently, the synergistic regulation of oxidative catalysis and proton conduction is achieved, which substantially accelerates the reaction kinetics of UOR.
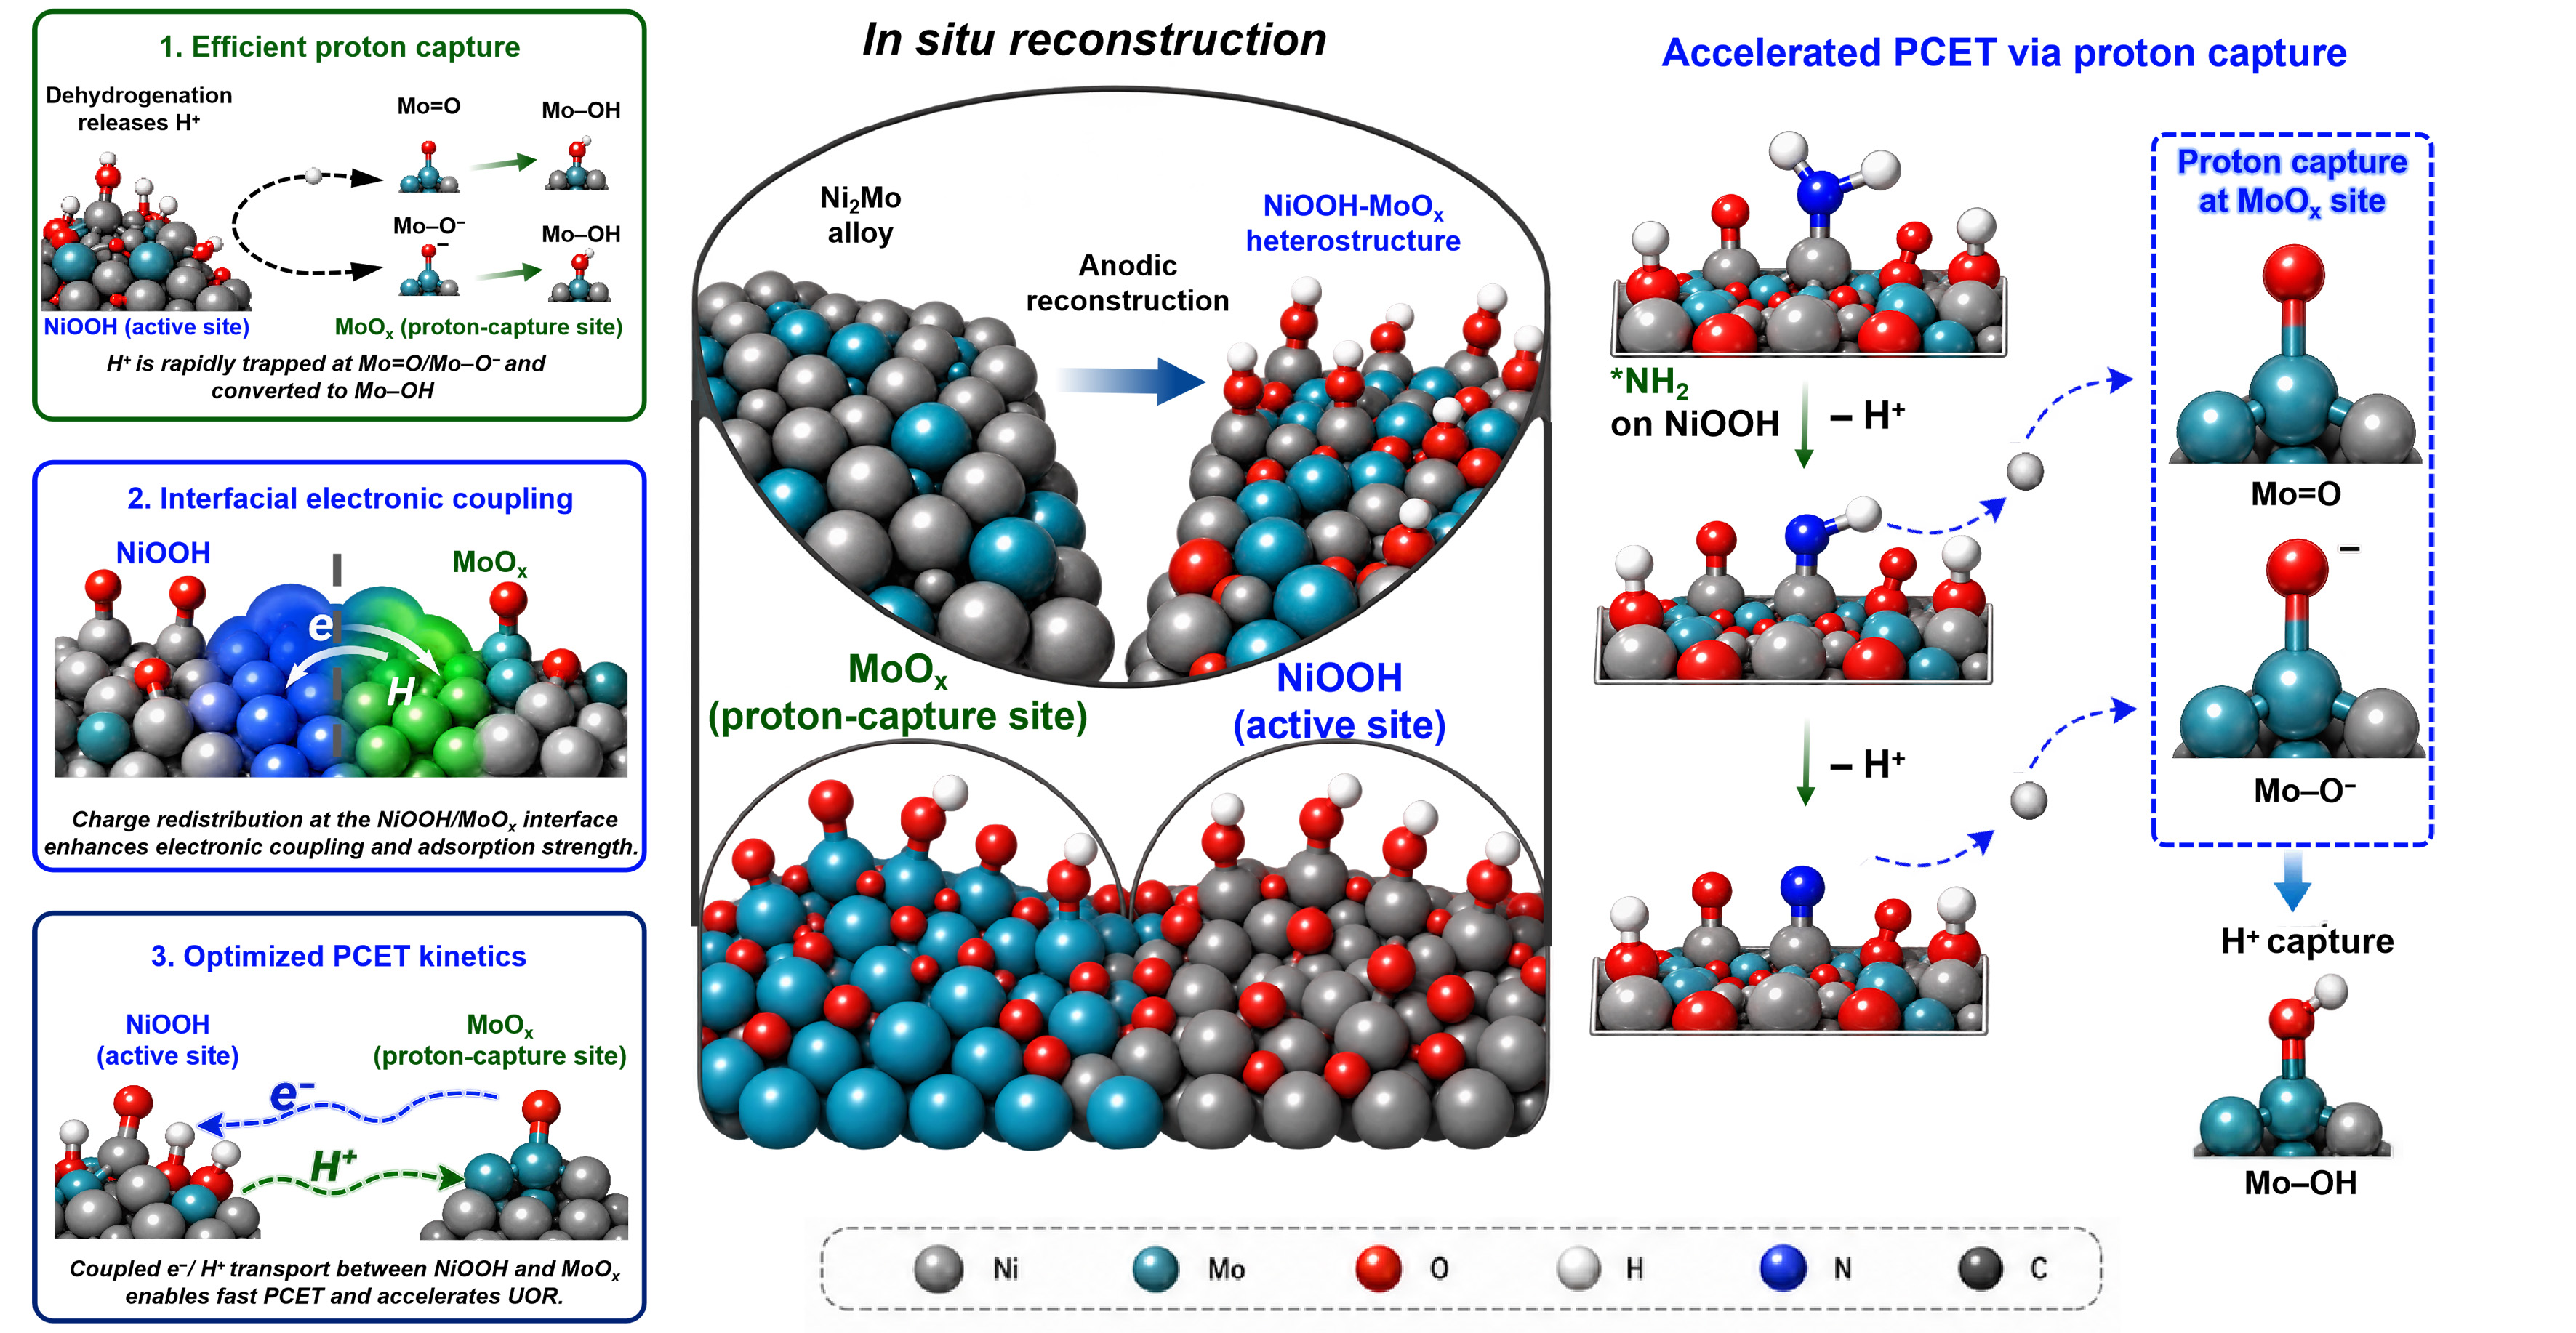
Figure 9. Schematic mechanism of the Ni2Mo-NiOOH/MoOx catalytic electrode during UOR. PCET: Proton-coupled electron transfer; UOR: urea oxidation reaction.
CONCLUSION
In summary, Ni2Mo-NiOOH/MoOx electrocatalyst was successfully fabricated via an anodic in situ reconstruction strategy, which exhibits outstanding UOR electrocatalytic activity and long-term structural stability. Benefiting from the significant enhancement of UOR catalytic activity induced by in situ reconstruction, the catalyst achieves a current density of 100 mA·cm-2 at only 1.341 V vs. RHE, which is merely 111 mV higher than the theoretical OER potential (1.23 V) and a small Tafel slope of 32.36 mV·dec-1, demonstrating favorable reaction kinetics. Structural characterization and electrochemical measurements confirm that the anodic reconstruction process not only transforms the Ni2Mo/NF into a Ni2Mo-NiOOH/MoOx composite structure, but also creates efficient proton-capturing sites rich in Mo=O/Mo-O- species. These sites can rapidly trap H+ released from the *-NH2 dehydrogenation process during UOR, effectively alleviate interfacial proton accumulation, and greatly lower the energy barrier of the key PCET step. Meanwhile, the Mo-induced local charge rearrangement further optimizes the adsorption/desorption energy barrier of UOR intermediates and improves the intrinsic catalytic activity of Ni active sites. In situ Raman spectroscopy, in situ EIS and electrolyte control experiments further verify that the in situ generated MoOx proton-capturing sites act as the core motive for accelerating reaction kinetics. The reversible proton transfer cycle of MoOx species synergizes with NiOOH active sites to construct efficient electron/proton synergistic transfer pathways for the UOR process. Attributed to the above superior performances, Ni2Mo-NiOOH/MoOx achieves a urea degradation efficiency of 97.21% within 420 min, and maintains a high current retention of 98.75% after continuous operation for 300 h at 100 mA·cm-2. It presents broad practical application prospects in the fields of urea electrolysis for hydrogen production and collaborative treatment of urea-containing wastewater. Furthermore, the proposed strategy of in situ reconstruction-mediated synergistic regulation of active sites and proton-capturing sites provides a novel design philosophy and universal reference for the development of high-efficiency UOR electrocatalysts.
DECLARATIONS
Authors’ contributions
Writing-original draft, methodology, investigation, formal analysis, data curation: Liu, J.
Writing-review & editing: Zhou, M.
Visualization, investigation: Wang, C.
Project administration: Zhou, L.
Conceptualization: Ruan, M.
Writing-review & editing, supervision, funding acquisition: Liu, Z.
Availability of data and materials
The original contributions presented in this study are included in the article/Supplementary Materials. Further inquiries can be directed to the corresponding authors.
AI and AI-assisted tools statement
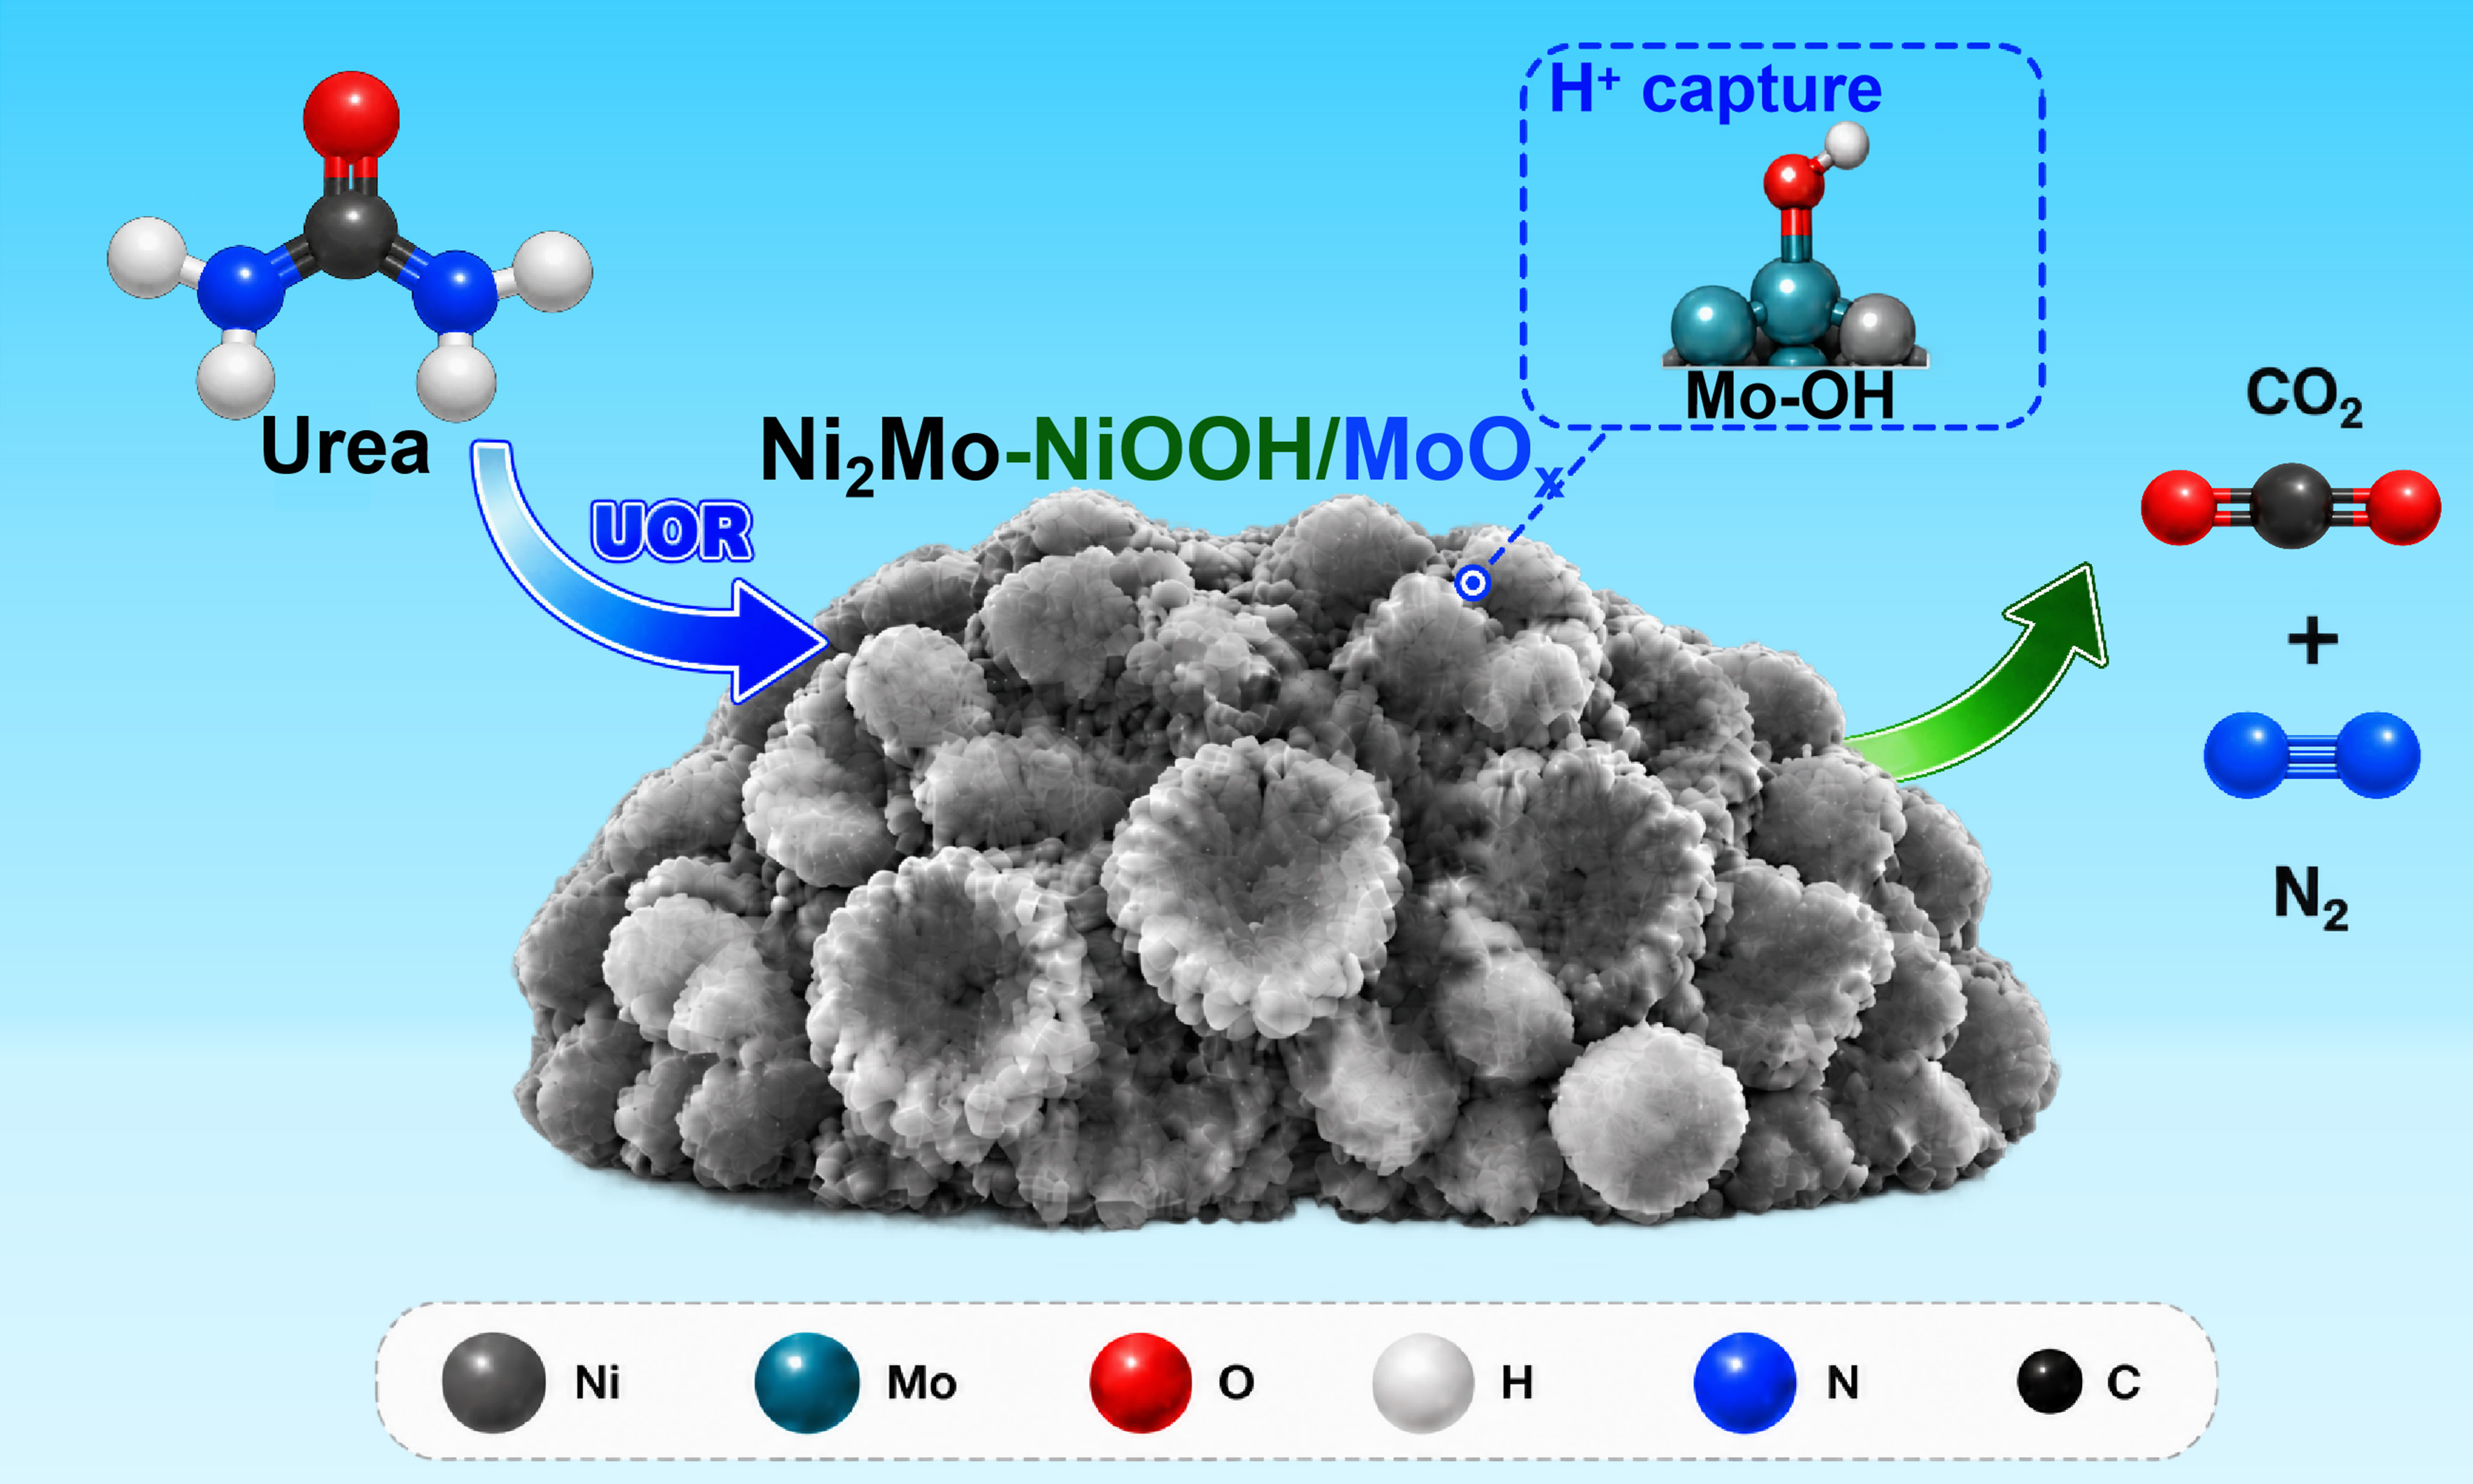
During the preparation of this manuscript, the AI tool WPS AI (WPS AI 1.0, algorithm filing identifier: Beijing-WPS-AI, dated 2023-10-16) was used for visual processing and graphical optimization of Figure 9 and the Graphical Abstract. Specifically, in Figure 9, WPS AI was used to segment/cut out the molecular structures, followed by their integration, layout adjustment, annotation, and final graphical optimization using WPS and WPS AI. In the Graphical Abstract, WPS AI was used to cut out and enhance the scanning electron microscopy (SEM) image of the sample acquired by the authors, followed by integration, layout adjustment, annotation, and final graphical optimization using WPS and WPS AI. The tool did not influence the study design, data collection, analysis, interpretation, or the scientific content of the work. All authors take full responsibility for the accuracy, integrity, and final content of the manuscript.
Financial support and sponsorship
The authors gratefully acknowledge the support from the National Natural the Science Foundation of China (No. 52373301, No. 22678414 and No. 22508289), the Science Foundation of Tianjin (No. 23JCZDJC00180 and No. 25JCQNJC00260), and the Joint Research Center of China and Foreign Countries Special Fund of Tianjin Innovation Platform (No. 24PTLYHZ00240).
Conflicts of interest
All authors declared that there are no conflicts of interest.
Ethical approval and consent to participate
Not applicable.
Consent for publication
Not applicable.
Copyright
© The Author(s) 2026.
Supplementary Materials
REFERENCES
1. Kempler, P. A.; Coridan, R. H.; Luo, L. Gas evolution in water electrolysis. Chem. Rev. 2024, 124, 10964-1007.
2. Zhang, W.; Liu, M.; Gu, X.; Shi, Y.; Deng, Z.; Cai, N. Water electrolysis toward elevated temperature: advances, challenges and frontiers. Chem. Rev. 2023, 123, 7119-92.
3. Yu, L.; Ning, M.; Wang, Y.; Yuan, C.; Ren, Z. Direct seawater electrolysis for hydrogen production. Nat. Rev. Mater. 2025, 10, 857-73.
4. Esfandiari, N.; Aliofkhazraei, M.; Colli, A. N.; et al. Metal-based cathodes for hydrogen production by alkaline water electrolysis: review of materials, degradation mechanism, and durability tests. Prog. Mater. Sci. 2024, 144, 101254.
5. dos Santos, K. G.; Eckert, C. T.; De Rossi, E.; et al. Hydrogen production in the electrolysis of water in Brazil, a review. Renew. Sustain. Energy Rev. 2017, 68, 563-71.
6. Yang, J.; Lam, T. Y.; Luo, Z.; Cheng, Q.; Wang, G.; Yao, H. Renewable energy driven electrolysis of water for hydrogen production, storage, and transportation. Renew. Sustain. Energy Rev. 2025, 218, 115804.
7. Yu, J.; Li, Z.; Wang, C.; et al. Engineering advanced noble-metal-free electrocatalysts for energy-saving hydrogen production from alkaline water via urea electrolysis. J. Colloid Interface Sci. 2024, 661, 629-61.
8. Zhang, L.; Wang, L.; Lin, H.; et al. A lattice-oxygen-involved reaction pathway to boost urea oxidation. Angew. Chem. 2019, 131, 16976-81.
9. Kong, Z.; Wu, J.; Liu, Z.; Yan, D.; Wu, Z. P.; Zhong, C. J. Advanced electrocatalysts for fuel cells: evolution of active sites and synergistic properties of catalysts and carrier materials. Exploration 2024, 5, 20230052.
10. Zemtsova, V. M.; Oshchepkov, A. G.; Savinova, E. R. Unveiling the role of iron in the nickel-catalyzed urea oxidation reaction. ACS Catal. 2023, 13, 13466-73.
11. Rasal, A. S.; Chen, H. M.; Yu, W. Electronic structure engineering of electrocatalyst for efficient urea oxidation reaction. Nano Energy 2024, 121, 109183.
12. Wang, L.; Zhu, Y.; Wen, Y.; et al. Regulating the local charge distribution of Ni active sites for the urea oxidation reaction. Angew. Chem. Int. Ed. Engl. 2021, 60, 10577-82.
13. Lee, J. W.; Lee, D.; Lee, H.; Shim, S.; Kim, J. H.; Lee, C. Enhanced oxidation of urea by pH swing during chlorination: pH-dependent reaction mechanism. Water Res. 2023, 242, 120183.
14. Zhou, L.; Feng, D.; Liu, C.; Sun, Y.; Fu, Y.; Ma, T. Amorphous Ni(OH)2-Ni3S2/NF nano-flower heterostructure catalyst promotes efficient urea assisted overall water splitting. Chem. Asian J. 2024, 19, e202300980.
15. Li, M.; Wu, X.; Liu, K.; et al. Nitrogen vacancies enriched Ce-doped Ni3N hierarchical nanosheets triggering highly-efficient urea oxidation reaction in urea-assisted energy-saving electrolysis. J. Energy Chem. 2022, 69, 506-15.
16. Zhang, J.; Song, X.; Kang, L.; et al. Stabilizing efficient structures of superwetting electrocatalysts for enhanced urea oxidation reactions. Chem. Catal. 2022, 2, 3254-70.
17. Zeng, F.; Mebrahtu, C.; Liao, L.; Beine, A. K.; Palkovits, R. Stability and deactivation of OER electrocatalysts: a review. J. Energy Chem. 2022, 69, 301-29.
18. Zheng, Z.; Wu, D.; Chen, L.; et al. Collaborative optimization of thermodynamic and kinetic for Ni-based hydroxides in electrocatalytic urea oxidation reaction. Appl. Catal. B Environ. 2024, 340, 123214.
19. Liu, Q.; Zhao, F.; Yang, X.; et al. Constructing interlaced network structure by grain boundary corrosion methods on CrCoNiFe alloy for high-performance oxygen evolution reaction and urea oxidation reaction. J. Mater. Sci. Technol. 2024, 203, 97-107.
20. Xu, S.; Jiao, D.; Ruan, X.; et al. O-2p hybridization enhanced transformation of active γ-NiOOH by chromium doping for efficient urea oxidation reaction. Adv. Funct. Mater. 2024, 34, 2401265.
21. Wang, Y.; Qian, G.; Xie, Z.; et al. Strengthening CuNiCoMo medium-entropy alloy by tuning local lattice tensile strain and built-in electric field for boosting urea oxidation reaction. Appl. Catal. B: Environ. Energy 2025, 365, 124841.
22. Du, M.; Ji, Y.; Li, Y.; (Frank) Liu, S.; Yan, J. Construction of an internal charge field: CoS1.097/Ni3S2 heterojunction promotes efficient urea oxidation reaction. Adv. Funct. Mater. 2024, 34, 2402776.
23. Ni, S.; Qu, H.; Xu, Z.; et al. Interfacial engineering of the NiSe2/FeSe2 p-p heterojunction for promoting oxygen evolution reaction and electrocatalytic urea oxidation. Appl. Catal. B Environ. 2021, 299, 120638.
24. Wang, P.; Zheng, W.; Qu, Y.; et al. Photo-excited high-spin state Ni (III) species in Mo-doped Ni3S2 for efficient urea oxidation reaction. Small 2024, 20, 2403107.
25. Xu, S.; Ruan, X.; Ganesan, M.; Wu, J.; Ravi, S. K.; Cui, X. Transition metal-based catalysts for urea oxidation reaction (UOR): catalyst design strategies, applications, and future perspectives. Adv. Funct. Mater. 2024, 34, 2313309.
26. Zhong, M.; Xu, M.; Ren, S.; et al. Modulating the electronic structure of Ni(OH)2 by coupling with low-content Pt for boosting the urea oxidation reaction enables significantly promoted energy-saving hydrogen production. Energy Environ. Sci. 2024, 17, 1984-96.
27. Zhang, S.; Jin, M.; Xu, H.; et al. An oxygen-coordinated cobalt single-atom electrocatalyst boosting urea and urea peroxide production. Energy Environ. Sci. 2024, 17, 1950-60.
28. Gao, X.; Zhang, S.; Wang, P.; Jaroniec, M.; Zheng, Y.; Qiao, S. Z. Urea catalytic oxidation for energy and environmental applications. Chem. Soc. Rev. 2024, 53, 1552-91.
29. Geng, S. K.; Zheng, Y.; Li, S. Q.; et al. Nickel ferrocyanide as a high-performance urea oxidation electrocatalyst. Nat. Energy 2021, 6, 904-12.
30. Zhang, J.; Zhu, J.; Kang, L.; et al. Balancing dynamic evolution of active sites for urea oxidation in practical scenarios. Energy Environ. Sci. 2023, 16, 6015-25.
31. Yang, C.; Pang, H.; Li, X.; et al. Scalable electrocatalytic urea wastewater treatment coupled with hydrogen production by regulating adsorption behavior of urea molecule. Nanomicro Lett. 2025, 17, 159.
32. Zheng, X.; Yang, J.; Li, P.; et al. Dual-atom support boosts nickel-catalyzed urea electrooxidation. Angew. Chem. 2023, 135, e202217449.
33. Liu, J.; Chen, Y.; Zhou, M.; Liu, Z. Synergistic Al-induced stabilization of Fe-O bonds to mitigate Fe3+ leaching and oxygen vacancy for enhanced urea oxidation catalysis. Chem. Eng. J. 2026, 531, 173953.
34. Lu, S.; Zheng, X.; Wang, H.; et al. Energy-saving hydrogen production by heteroatom modulations coupling urea electrooxidation. EcoMat 2024, 6, e12477.
35. Gao, X.; Bai, X.; Wang, P.; et al. Boosting urea electrooxidation on oxyanion-engineered nickel sites via inhibited water oxidation. Nat. Commun. 2023, 14, 5842.
36. Jiang, W.; Zhang, J.; Wu, J.; et al. Regulating the competitive adsorption of urea and OH− via brønsted base intercalated nickel sites for highly selective urea oxidation. Adv. Energy Mater. 2026, 16, e03978.
37. Yang, J.; Wang, Y.; Fan, Y.; Xu, B.; Tu, T. Sustainable recycling of polyamides via C-N bond cleavage facilitated by methyl tosylate under mild conditions. Nat. Commun. 2025, 16, 8884.
38. Ni, X.; Luo, Z.; Sun, H.; et al. Multi-site relay catalysis for efficient hydrogen production coupled with urea-rich wastewater treatment at large current density. Appl. Catal. B: Environ. Energy 2026, 399, 127132.
39. Ma, Z.; Liu, X.; Wang, H.; et al. Interfacial engineering of NiCo nanocones to promote NiOOH formation via enhanced dehydrogenation for efficient urea electrooxidation. Adv. Energy Mater. 2026, 16, e02563.
40. Chen, W.; Xu, L.; Zhu, X.; et al. Unveiling the electrooxidation of urea: intramolecular coupling of the N-N bond. Angew. Chem. Int. Ed. Engl. 2021, 60, 7297-307.
41. Sari, F. N. I.; Tran, N. T. T.; Ke, M.; Purba, Y.; Huynh, T. M. D.; Ting, J. M. Suppression of oxygen electrooxidation toward enhanced urea electrooxidation via balancing the adsorption of OH- and urea intermediates in quaternary-metal LDH. Chem. Eng. J. 2025, 515, 163966.
42. Wang, J.; Sun, M.; Zhang, X.; et al. pH-dependent urea electrooxidation: from mechanism to catalysts and applications. Adv. Mater. 2026, 38, e15043.
43. Gong, Z.; Xu, M.; Li, P.; Ye, G.; Li, W.; Fei, H. Rapid reconstruction of amorphous/crystalline heterojunction boosts nickel-catalyzed urea electrooxidation. ACS Catal. 2025, 15, 20958-67.
44. Yang, H.; Li, T.; Li, Z.; Huang, K.; Zhang, Z.; Meng, X. Dual-metal regulation of PCET and spin polarization synergism for efficient photocatalytic urea synthesis. Chem. Eng. J. 2025, 522, 167328.
45. Derosa, J.; Garrido-Barros, P.; Li, M.; Peters, J. C. Use of a PCET mediator enables a Ni-HER electrocatalyst to act as a hydride delivery agent. J. Am. Chem. Soc. 2022, 144, 20118-25.
46. Hou, L.; Sun, C.; Zhang, Z.; et al. Carbon-oxyanion atomically steering direct urea oxidation on NiOOH at industrial current densities. Adv. Funct. Mater. 2025, 36, e19865.
47. Wang, P.; Gao, X.; Zheng, M.; Jaroniec, M.; Zheng, Y.; Qiao, S. Z. Urine electrooxidation for energy-saving hydrogen generation. Nat. Commun. 2025, 16, 2424.
48. Yu, X.; Zhang, W.; Ma, L.; et al. Interfacial engineering of a CoSe@NiFe heterostructure electrocatalyst for high-efficiency water and urea oxidation. Green Chem. 2025, 27, 731-42.
49. Zeng, M.; Wu, J.; Li, Z.; et al. Interlayer effect in NiCo layered double hydroxide for promoted electrocatalytic urea oxidation. ACS Sustainable Chem. Eng. 2019, 7, 4777-83.
50. Liu, J.; Zhou, M.; Chen, Y.; Liang, X.; Liu, Z. Etching reconstruction of ultrathin layered NiFe-hydroxide: building synergistic VFe(III)/Ni(III) active structures for an efficient urea oxidation reaction. Green Chem. 2026, 28, 11533-47.
51. Wang, L.; Zhao, Z. Dynamic adsorption of molybdate promoted self-optimization and self-healing of NiFe/NiMo in alkaline water electrolysis. Small 2026, 22, e10965.
Cite This Article
 Open Access
How to Cite
Liu, J.; Zhou, M.; Wang, C.; Zhou, L.; Ruan, M.; Liu, Z. Unveiling proton-regulated Mo=O/Mo-O- centers in reconstructed Ni2Mo-NiOOH/MoOx interfaces toward accelerated proton-coupled electron transfer for urea oxidation reaction. Catal. Energy Environ. 2026, 1, 9. https://dx.doi.org/10.20517/cee.2026.18
Download Citation
Export Citation File:
Type of Import
Tips on Downloading Citation
Citation Manager File Format
Type of Import
Direct Import: When the Direct Import option is selected (the default state), a dialogue box will give you the option to Save or Open the downloaded citation data. Choosing Open will either launch your citation manager or give you a choice of applications with which to use the metadata. The Save option saves the file locally for later use.
Indirect Import: When the Indirect Import option is selected, the metadata is displayed and may be copied and pasted as needed.
About This Article
Copyright

Data & Comments
Data
0



Comments
Comments must be written in English. Spam, offensive content, impersonation, and private information will not be permitted. If any comment is reported and identified as inappropriate content by OAE staff, the comment will be removed without notice. If you have any queries or need any help, please contact us at support@oaepublish.com.